In organic reactions the catalyst is often the most expensive component and usually very toxic. Its incorporation into a solid support has been of great interest due to several advantages such as simplification of product work-up, separation, isolation, and reuse of the catalysts. In 1963, Merrifield introduced the concept of using supported species to overcome some of the physical and mechanical difficulties of completing long syntheses rapidly and efficiently.[ Reference Merrifield 1 ] Merrifield highlighted a solid-phase method for oligopeptide synthesis using polystyrene-based resins as a precursor. In this method, the syntheses of peptides is achieved by consecutive reactions involving supported protecting groups where the separation of the new supported species is not possible and hence the reactions must be clean and very high yielding. The use of polymer-supported reagents is less demanding. Over the last decade, polymer-supported catalysts have been extensively used in organic synthesis.[ Reference Madhavan, Jones and Weck 2 , Reference Clapham, Reger and Janda 3 ] By attaching the catalyst onto a polymer support it can be easily recovered after each reaction for recycling and not every site needs to be available for reaction.[ Reference Hodge 4 ] However, these immobilized catalysts are sometimes less active than the corresponding original catalysts and the precursors can be expensive and difficult to synthesize.[ Reference Sherrington 5 ]
During the last few years, the interest in developing novel polymeric nanostructures as scaffolds for catalysis has increased significantly.[ Reference Nuyken, Persigehl and Weberskirch 6 – Reference Chi, Scroggins and Frechet 8 ] Nanometer sized core-shell structured containers can be applied as catalytic nanoreactors where functionalities can be incorporated on the surface, the shell, or the well-defined core of the polymeric support. The advantages of such nanostructures over traditional supports are numerous since they combine benefits from both homogeneous and heterogeneous catalysts creating a novel environment for reaction and improved recovery.
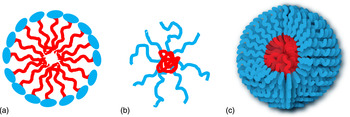
With the introduction of controlled radical polymerization (CRP) techniques the range of accessible polymer architectures, which have the capacity to act as nanoreactors for organic reactions, has now greatly increased.[ Reference Perrier and Takolpuckdee 9 – Reference Benoit, Chaplinski, Braslau and Hawker 11 ] Given the number of examples of catalytic polymeric nanoreactors in the literature, due to space limitations, in this prospective we will focus on core-shell-type polymeric nanoparticles, where the specific catalytic groups are tethered within the core domain and protected from degradation or side reactions by the surrounding corona. This allows for the selective reaction of the reagents within the central domain in an overall homogeneous media (Fig. 1). A few selected examples are discussed illustrating the benefits of tethering the catalyst in the core of the polymeric nanoreactor.
Different types of micelles and micelle analogous structures: (a) small molecule surfactants, (b) star-like polymer, (c) amphiphilic BCP micelle. Red domain is hydrophobic and blue domain is hydrophilic.

Well-defined polymeric nanostructures
To obtain well-defined nanoreactors, constituent polymers must have a low polydispersity (PDI) and a well-defined molecular weight. Moreover, the selected polymerization technique must be tolerant to functional groups and allow for the tuning of the structure and density of catalyst along the polymer chain. Living polymerization techniques, such as anionic or cationic procedures, are popular methods for synthesizing block copolymers (BCPs).[ Reference Szwarc 12 ] The polymer scaffold can be made in stages, each stage containing a different monomer and hence different characteristics (functionalities) can be introduced in each block.[ Reference Thurmond, Kowalewski and Wooley 13 ] However, ionic polymerizations are not tolerant to a broad range of functionalities and often non-trivial protecting chemistries are needed to prepare functional BCPs. Furthermore, the methodology is unsuitable for large-scale industrial applications due to the extremely stringent reaction conditions and high monomer purity, hence conventional radical chemistry is often employed in industry. Although radical techniques avoid the need for stringent reaction conditions, they lack the high level of control which a living polymerization offers.
The synthesis of a wide range of precisely engineered polymers has been achieved over the last two decades with the development of CRP techniques that share a lot of the features of living polymerizations.[ Reference Matyjaszewski and Tsarevsky 14 – Reference Bosman, Vestberg, Heumann, Fréchet and Hawker 16 ] Polydispersities obtained for CRPs can approach the values typically obtained for living procedures (ca. 1.1)[ Reference Hawker 17 ] and are often tolerant to trace impurities in the reaction medium. CRPs are based on the establishment of a dynamic equilibrium in solution between active (propagating) and deactivated (dormant) species. The dynamic equilibrium can be achieved by two general mechanistic approaches: the first one involves the reversible deactivation of propagating chains that subsequently can be reactivated, either catalytically as in atom transfer radical polymerization (ATRP)[ Reference Wang and Matyjaszewski 18 ] or spontaneously like in nitroxide-mediated polymerization (NMP).[ Reference Hawker, Barclay, Orellana, Dao and Devonport 19 ] The second approach relies on the degenerative transfer between propagating chains and dormant species with a typical example of this being reversible addition-fragmentation chain transfer (RAFT) polymerization.[ Reference Chiefari, Chong, Ercole, Krstina, Jeffery, Le, Mayadunne, Meijs, Moad, Moad, Rizzardo and Thang 20 ] One of the main advantages of CRPs is that they can tolerate a wide range of functionalities while retaining excellent control over the molecular characteristics of the products and hence allowing the synthesis of precisely engineered BCP macromolecular architectures. These features give the ability to readily tune the chemical nanoenvironment of a given functionality with excellent precision within the BCP.[ Reference Braunecker and Matyjaszewski 21 ]
Hydrophobic pockets for organic synthesis in aqueous media
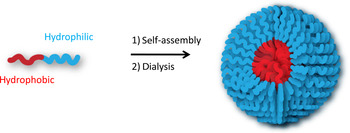
The use of water as a reaction medium has experienced a great development in the last few decades.[ Reference Sheldon 22 , Reference Gruttadauria, Giacalone and Noto 23 ] Despite water being cheap, safe, and the most abundant solvent on earth, its application in organic synthesis is restricted by the low solubility of organic compounds in water and the decomposition or deactivation of some substrates in aqueous conditions. A good approach to overcome these challenges has been the introduction of surfactants and water-soluble dendrimers as reaction pockets.[ Reference Vriezema, Aragones, Elemans, Cornelissen, Rowan and Nolte 24 , Reference Helms and Fréchet 25 ] By carrying out organic reactions in the presence of surfactant-type micelles, enhancement of rates and/or specificity has been observed for different reactions based on hydrophobic forces. The high concentration created in the hydrophobic core enhances the reaction rates and induces selectivity in the same way as enzymes in biological systems. Unfortunately, problems with catalyst recovery and product isolation limit the scope of this approach. In the same way as surfactant-based micellar systems, polymeric micelles can concentrate reactants within a well-confined nanoenvironment. By carefully tuning the nature of the monomeric units of the core block, the selectivity of the reaction can be modified based on non-covalent interactions core-substrates such hydrogen-bonding or solvophobic effects.[ Reference McHale, Patterson, Zetterlund and O'Reilly 26 , Reference Cotanda and O'Reilly 27 ] A further advantage of polymeric self-assembled systems is their stability over surfactant-based micelles, creating a more “protected” kinetically frozen pocket within the core domain.[ Reference Cotanda and O'Reilly 27 – Reference Arumugam, Vutukuri, Thayumanavan and Ramamurthy 29 ] Moreover, by covalently attaching the catalytic functionality within the polymer backbone, it can be easily recovered after reaction by precipitation or ultrafiltration methods.[ Reference Liu, Wang, Wang, Lu, Piñón and Weck 30 , Reference Cotanda, Lu, Patterson, Petzetakis and O'Reilly 31 ] Therefore, polymeric core-shell-type catalytic nanoreactors have been investigated in which the hydrophobic core provides a favorable confined environment for the hydrophobic starting materials, while the hydrophilic shell guarantees water solubility. The simplest method for the assembly of synthetic polymer chains into nanoparticles is the solution self-assembly to allow for the formation of spherical polymer micelles. Conventional micelles based on hydrophilic-hydrophobic diBCPs have been extensively reported.[ Reference Wooley 32 ] The major driving force behind the self-assembly of amphiphilic copolymers is the decrease in free energy of the system due to the removal of the hydrophobic fragments from the incompatible aqueous environment. The micelle core is stabilized and “protected” from the surrounding aqueous media by the hydrophilic blocks. Amphiphilic micelles are formed spontaneously via the solution-state self-assembly of amphiphilic (multi)-BCPs, consisting of hydrophobic and hydrophilic chain segments (Fig. 2). The self-assembly process is usually achieved by the so-called solvent switch method, which involves dissolving the amphiphilic copolymer in a good solvent for all the blocks followed by the gradual addition of a non-solvent for one of the blocks (selective solvent) at a concentration above the critical micelle concentration (cmc). Then via dialysis or evaporation, the complete replacement of the good solvent by the non-solvent is achieved and well-defined aggregates (micelles) are prepared. Furthermore, depending on the combination of polymer-solvent, the final system can be kinetically trapped (frozen micelles), where no unimer exchange is observed or dynamic, where fast intermicellar polymer chain exchange is allowed and the solvent, to a certain extent, penetrates the solvophobic core.[ Reference Nicolai, Colombani and Chassenieux 33 ]
Self-assembled spherical micelles from amphiphilic BCPs in water.

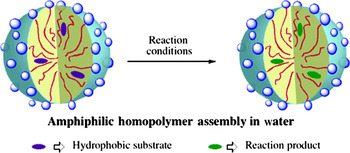
The kinetically frozen nature of polymeric micelles in water creates a confined hydrophobic pocket, which avoids water permeability creating a stable environment for substrates sensitive to hydrolysis. Thayumanavan and Ramamurthy recently reported a comparative study of product selectivity using amphiphilic polymer versus surfactant based micelles in water (Fig. 3).[ Reference Arumugam, Vutukuri, Thayumanavan and Ramamurthy 29 ] Their studies revealed that the hydrophobic domains created by a kinetically frozen styrenic-based amphiphilic homopolymer stops hydrolysis of hydrophobic substrates and offers better selectivity control than micelles formed from small molecule surfactants. This is due to the more stable and confined hydrophobic pocket generated by the amphiphilic homopolymer compared with that for small molecule surfactants.
Schematic representation of a reaction within the homopolymer micelle-type nanocontainer. Reproduced with permission from Ref. Reference Arumugam, Vutukuri, Thayumanavan and Ramamurthy29. Copyright 2005 American Chemical Society.

Functional amphiphilic BCPs have been self-assembled into micelles in water with incorporation of functional groups within the core, enabling different reactions to be carried out within this unique hydrophobic environment. Nuyken and co-workers pioneered the development of a platform of amphiphilic, water-soluble BCPs for metal catalysis in water.[ Reference Nuyken, Persigehl and Weberskirch 6 ] The covalent immobilization of transition metal catalysts (Rh, Ir, or Pd) within the hydrophobic core increased the local concentration of the catalyst in the reaction.[ Reference Gall, Bortenschlager, Nuyken and Weberskirch 34 , Reference Persigehl, Jordan and Nuyken 35 ] The hydrophobic substrates were dissolved in the micellar core, where the catalyst was also located, allowing the transformations to be carried out in an overall aqueous environment, significantly increasing the reaction rates compared with the water-soluble homopolymer counterparts.[ Reference Zarka, Nuyken and Weberskirch 36 ] However, the leaching of the catalyst could not be avoided therefore the recyclability of this polymeric systems was limited.
In further work by Nuyken's, unfunctionalized polymers were synthesized by living cationic polymerization of 2-oxazoline monomers and the active catalyst was introduced via a post-functionalization method. While this approach affords well-defined precursor polymers the post-modification method can often be inefficient. Hence, a new strategy has been developed which involves the synthesis of functionalized monomers, which can be incorporated selectively into a specific domain of a BCP via robust CRP techniques.[ Reference Liu, Wang, Wang, Lu, Piñón and Weck 30 , Reference Ievins, Wang, Moughton, Skey and O'Reilly 37 – Reference O'Lenick, Jiang and Zhao 40 ] This approach allows for the synthesis of functional polymers without the need of protecting groups or post-functionalization steps.[ Reference Lu, Smart, Epps, Longbottom and O'Reilly 41 ] The choice of functional monomer provides the possibility of introducing a variety of functional groups into the micelle (core, shell, or surface), which can tune the properties of the nanostructure for different applications. By introducing a stimuli-responsive block in the synthesis of block co-polymers, one block can be tunable to be hydrophilic or hydrophobic by applying an external stimulus, whereas the other block remains hydrophobic or hydrophilic offering convenient ways to control the self-assembly. Following this strategy, the concept of micelles as pseudohomogeneous catalyst supports has been explored using recoverable and responsive polymers.[ Reference Ge, Xie, Chen, Jiang, Zhang, Liu and Liu 39 , Reference Wang, Wei, Zhang, Jiang, Zheng, Shi and Dong 42 ]
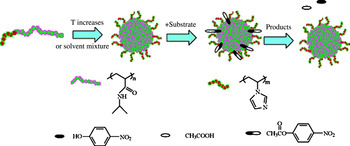
Most significantly, Liu and co-workers used RAFT to synthesize stimuli-responsive BCPs, poly(N-isopropylacrylamide)-b-poly(N-vinylimidazole) (PNIPAM-b-PVim), which self-assembled above the lower critical solution temperature (LCST) of PNIPAM into micelles with the hydrophobic PNIPAM core of the micelles.[ Reference Ge, Xie, Chen, Jiang, Zhang, Liu and Liu 39 ] Interestingly, the authors in this case incorporated the catalytic PVim functionality in the shell of the aggregates and studied the hydrolysis reaction of p-nitrophenyl acetate (NPA) at elevated temperatures, where the BCP is above its critical micellization temperature (CMT) and thus micelles were formed (Fig. 4). They found that in the presence of the catalytic micelles, the rate of esterolysis increased up to 7 times, and they proposed that this was due to the hydrophobic affinity of the reactants for the hydrophobic PNIPAM core bringing the reactants in close proximity to the catalytic shell and hence increased their local concentration. Although in this example the authors left the catalyst “exposed’’ in the shell domain and the reaction did not take place in the core of the aggregates, the importance of selective localization of the starting materials via hydrophobic attractions was illustrated.
Stimuli-responsive double hydrophilic BCP micelles with switchable catalytic activity. Reproduced with permission from Ref. Reference Ge, Xie, Chen, Jiang, Zhang, Liu and Liu39. Copyright 2007 American Chemical Society.

O'Reilly et al. recently reported the successful incorporation of the organocatalyst l-proline into the hydrophobic segment of two amphiphilic BCPs via RAFT giving two efficient nanoreactor systems for aldol reactions in water.[ Reference Lu, Cotanda, Patterson, Longbottom and O'Reilly 28 ] The catalytic efficiency of these core–shell structures was investigated using the reaction of 4-nitrobenzaldehyde and cyclohexanone. The polymeric nanoreactors showed excellent catalytic properties, resulting in higher activity in water compared with unsupported l-proline in water and organic solvents. Moreover, it was proved that the different hydrophobicity of the micellar core determined the amount of water present in the reaction environment affecting the reaction rates.
In 2009, Zhao and co-workers used ATRP to synthesize a poly(ethylene oxide)-b-poly(methoxydi (ethylene glycol) methacrylate-co-2-(N-methyl-N-(4-pyridyl)amino)ethyl methacrylate), (PEO-b-P(DEGMMA-co-MAPMA)) diBCP.[ Reference O'Lenick, Jiang and Zhao 40 ] In this case MAPMA, a 4-dimethylaminopyridine (DMAP) functionalized monomer, was copolymerized with the temperature-responsive DEGMMA monomer. The corresponding temperature-responsive block with the tethered DMAP functionality formed the hydrophobic core of the micelle upon heating to 30-48 °C.[ Reference O'Lenick, Jiang and Zhao 40 ] They envisioned that, by increasing the temperature above the LCST of PDEGMMA, hydrophobic substrates could be concentrated in the core of the micelle by hydrophobic attractions and proposed that this would result in an enhanced reaction rate due to the high concentration of substrates within the catalytic core domain. To explore this, the authors studied the hydrolysis of NPA at different temperatures, always above the CMT of the aggregates. Disappointingly, the reaction rates did not increase with temperature as much as expected by the Arrhenius equation and the result was attributed to a poor partitioning of the starting materials to the hydrophobic core.
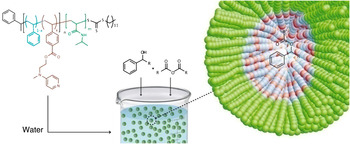
Recently our group utilized DMAP-functionalized BCP micelles to effectively catalyze acylation reactions in water.[ Reference Cotanda, Lu, Patterson, Petzetakis and O'Reilly 31 ] We investigated the effect of having a DMAP functionality covalently attached to the polystyrene core of polymeric micelles synthesized by RAFT polymerization (Fig. 5). The reactivity of the tethered organocatalyst within the nanostructure was found to be high, improving in some cases the acylation rates up to 100 times compared with those for unsupported DMAP in organic solvents. The efficacy of the system was affected by the hydrophobicity of the starting materials; the more hydrophobic substrates exhibited more favorable interactions with the hydrophobic polystyrene core and hence greater partitioning was observed. This effect is directly related to the selectivity of the nanoreactors toward more hydrophobic substrates. Moreover, as we designed our BCPs to contain a temperature responsive PNIPAM shell, the catalytic nanoreactors were easily recycled by precipitation with a simple heating step. Several recycling steps were carried out without any side effects on the catalytic performance of the DMAP nanoreactors.
DMAP nanoreactor for efficient catalysis of hydrophobic substrates in water. Reproduced with permission from Ref. Reference Cotanda, Lu, Patterson, Petzetakis and O'Reilly31. Copyright 2012 American Chemical Society.

Based on these initial results, the concept of substrate recognition based on hydrophobic interactions was further explored.[ Reference Cotanda and O'Reilly 27 ] We showed that these polymeric organocatalytic nanoreactors are not only very efficient and recyclable supports for catalytic reactions in aqueous media, but also show specific substrate recognition able to drastically modify the selectivity of the reactions based on the simple concept of hydrophobicity. The hydrophobic core-substrate attraction induced by the unique nature of the polymeric micelle creates a concentrated catalytic environment that allows for reaction of otherwise nonreactive species significantly improving the effects observed for surfactant-based systems. This is the first example of polymeric nanoreactors capable of effectively distinguishing from a pool of substrates of similar or different reactivities and where specificity is achieved based on substrate hydrophobicity. We see these results as an initial approach toward enzyme mimics for organic reactions in water.
Cross-linked polymeric micelles
Although polymeric micelles can be more stable than surfactant-based micelles, these structures can be affected by drastic changes in concentration or temperature. The stabilization of polymeric micelles through cross-linking, to obtain stable robust particles, has been extensively reported in the literature. Two different techniques are typically used: one involves the formation of the aggregates followed by the addition of a cross-linking agent that can specifically crosslink the core, shell, or corona.[ Reference Wooley 32 , Reference O'Reilly, Hawker and Wooley 43 ] The second method involves the preparation of polymeric micelles using BCPs containing self-crosslinkable groups, and then inducing the post-modification cross-linking process.[ Reference Yan, Liu, Liu, Tang, Peng, Pakhomov and Wong 44 , Reference Liu, Pinon and Weck 45 ] Surprisingly, only a few examples of their application in catalysis have been reported.[ Reference O'Reilly, Hawker and Wooley 43 ]
Catalytic shell cross-linked micelles have been synthesized by our group using NMP. In this case, the core of the particles contained a terpyridine side group that could be utilized as a ligand to bind Cu(I) species.[ Reference Ievins, Wang, Moughton, Skey and O'Reilly 37 ] The catalytic efficiency of these nanoparticles on mediating “click’’ reactions was confirmed using a fluorogenic reaction between an azido coumarin and an alkynyl small molecule. Further, more recent work by Weck and co-workers used poly(2-oxazoline)-based amphiphilic triBCPs with a salen-functionalized hydrophobic block prepared by living cationic polymerization.[ Reference Liu, Wang, Wang, Lu, Piñón and Weck 30 ] The polymers were self-assembled into core-shell-corona micelles and stabilized by photo-crosslinking the mid-block. Subsequently, the catalytic sites were generated by adding Co(III)-producing Co(III)-salen complexes in the hydrophobic core of the particles. The authors explored the kinetic resolution of a range of epoxides of different hydrophobicities. These nanoreactors also exhibited substrate selectivity based on hydrophobic effects. However, at the same catalyst loading, the non-cross-linked catalyst showed a slightly higher catalytic activity due to the less permeable shell in the cross-linked particles. The same effect has been recently observed by Stenzel et al. when comparing the rate of release for a drug encapsulated in the core of a cross-linked and non-cross-linked micelle. Despite its high stability, the shell-cross-linked micelles lead to a network formation, which can act as obstacle for the diffusion drugs, leading to a reduced release rate.
Site isolation
One-pot multistep reactions are effective at reducing the waste and the cost of a synthetic process because they decrease the number of work-ups and purification steps, as well as the volume of the solvent used. These reactions are especially useful when multiple catalysts are used so that one can trap an unstable intermediate formed by the other. Although a variety of these reactions have been reported, they are limited to a relatively small number of systems where the catalysts are compatible with each other.[ Reference Broadwater, Roth, Price, Kobaslija and McQuade 46 , Reference Tietze 47 ] The work of Patchornik in 1981 demonstrated that this limitation can be overcome by immobilizing incompatible catalysts on solid supports.[ Reference Cohen, Kraus and Patchornik 48 ] The principle of site isolation to catalysis has been applied making use of different supports. Several examples show how polymeric nanostructures render incompatible catalysts compatible. In recent years, soluble dendritic and other hyperbranched polymers have emerged as attractive systems for the encapsulation and isolation of various functional groups within the interior of the polymers. The placement of catalytic functionality at the interior of a globular dendritic structure protects the catalyst from deactivation and allows for the tuning of molecular properties and catalytic activity by modification of the periphery and the interior environment, respectively.[ Reference Helms and Fréchet 25 ] Such stable unimolecular supports can exist with high stability in different solvents and temperatures; however, their synthetic difficulty limits their use.
Cascade reactions in self-assembled polymeric nanoreactors have been explored by Weberskirch et al. for the mono (Rh) and bimetallic (Rh and Ir) micellar catalysis in the hydroaminomethylation of 1-octene. Unfortunately, conditions optimized for one reaction were not favorable for the next one, probably due to the competition of more than one substrate for the metal center and the deactivation of the catalyst.[ Reference Gall, Bortenschlager, Nuyken and Weberskirch 34 ]
Polymeric stars
A similar but simpler approach to obtain stable polymeric nanoreactors is to synthesize unimolecular micelles. The facile synthesis and tunable composition of star polymers makes them an attractive alternative to the widely studied dendritic systems or other cross-linked micelles. A polymeric star is a macromolecule containing a single branched core with multiple linear chains or arms forming a 3D globule. The use of CRP techniques allows the formation of unimolecular branched stars and the selection of initiators monomers or cross-linkers allows for covalent incorporation of functionalities into the structure. The synthesis of polymeric stars is commonly carried out via three different strategies based on the sequence of formation of the core and the arms. In the “core-first” approach, polymeric arms are grown from a multifunctional cross-linked initiator, while in the “arm-first” approach presynthesized linear polymers are cross-linked using divinyl compounds.[ Reference Matyjaszewski and Tsarevsky 14 – Reference Bosman, Vestberg, Heumann, Fréchet and Hawker 16 , Reference Gao 49 ] A third strategy, “coupling onto”, consist of attaching linear polymers onto a multifunctional core. The “arm-first” method is the most commonly used strategy to synthesize stars by the most popular CRP techniques (NMP, RAFT, and ATRP).
Although it has already been demonstrated that cross-linking the core or the shell domain reduces the micelle permeability, the introduction of catalytic motif into the core of a polymeric star is a simple way of protecting the catalyst from the surrounding environment and avoid side reactions. Such structures share the recyclability of cross-linked micelles and have shown outstanding properties for site isolation.
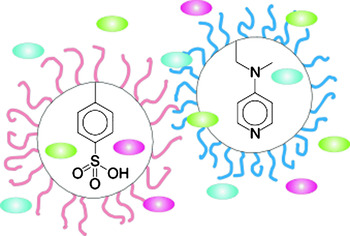
The groups of Fréchet and Hawker have developed an effective method to synthesize star polymers with core confinement of functionalities by NMP using the arm first approach. In 2005, Fréchet et al. report the synthetic utility of site isolation by carrying out a one pot cascade reaction of the otherwise incompatible acid and base catalysts (Fig. 6).[ Reference Helms, Guillaudeu, Xie, McMurdo, Hawker and Frechet 50 ] They synthesized star polymers that contained acid (para-toluenesulfonic acid) or base (DMAP) catalysts in their cores to perform acid and base cascade reactions in the same pot with an acid-catalyzed deprotection followed by the nucleophilic amine-catalyzed Baylis-Hillman reaction. These capsules were recovered after reaction and yielded higher activities in acylation reactions than linear polymers or other solid supports.
One-pot-reaction cascade involving sequential acid-catalyzed acetal hydrolysis followed by the amine-catalyzed Baylis-Hillman reaction.[ Reference Helms, Guillaudeu, Xie, McMurdo, Hawker and Frechet 50 ] Reproduced with permission from Ref. Reference Helms, Guillaudeu, Xie, McMurdo, Hawker and Frechet50. Copyright 2005 Wiley Interscience.

This ability to generate an otherwise incompatible cascade of reactions is characteristic of biological systems where different enzymes are used consecutively to create a wide range of chemical transformations through the combination of simple steps. Further studies reported that site isolation with star polymers enables the combination of incompatible catalysts for more sophisticated asymmetric cascade reactions. This group have also published the design of non-interpenetrating star polymers to combine iminium, enamine, and hydrogen bond catalysis in one pot for asymmetric reactions that generate cascade products with more than one chiral center.[ Reference Chi, Scroggins and Frechet 8 ]
The groups of Sawamoto and Matyjaszewski have extensively reported the formation of functional polymeric stars by ATRP.[ Reference Baek, Kamigaito and Sawamoto 51 – Reference Terashima, Ouchi, Ando and Sawamoto 53 ]
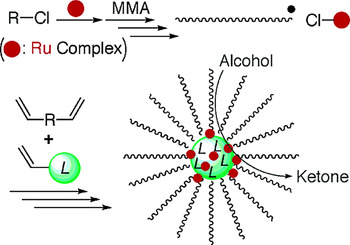
In 2003, the group of Sawamoto reported an interesting procedure to synthesize catalytic stars where a ruthenium complex was incorporated into the core using ruthenium-catalyzed living radical polymerization.[ Reference Terashima, Kamigaito, Baek, Ando and Sawamoto 52 ] The ruthenium catalyst was encapsulated by cross-linking poly (methylmethacrylate) “arms” using ethylene glycol dimethylacrylate in the presence of diphenyl-4-styrylphosphine as a ligand to form the catalytic core. The ruthenium-containing star polymers were then employed as a catalyst for the oxidation of alcohols into ketones (Fig. 7). However, they observed that the activity of the stars was lower than the unsupported complex in solution, probably due to the overloading of catalyst in the core of the star. Sawamoto and co-workers cleverly used a predicted “drawback” of ATRP (difficult removal of metal catalyst) to synthesize a very efficient and highly recyclable nanoreactor. In this case, the residual metal catalyst used to synthesize the nanoreactor by ATRP and was encapsulated and employed as a catalyst for organic reactions. Furthermore, the same star polymer catalysts showed high activity, versatility, functionality tolerance, and recyclability as a catalyst for ATRP reactions.[ Reference Terashima, Nomura, Ito, Ouchi and Sawamoto 54 ]
Direct encapsulation of metal catalyst into star polymer core during metal-catalyzed living radical polymerization. Reproduced with permission from Ref. Reference Terashima, Kamigaito, Baek, Ando and Sawamoto52. Copyright 2003 American Chemical Society.

Conclusions
Although supported catalysis has been a popular subject for many years, the study of polymeric nanostructures for catalysis of organic reactions has received increasing interest in the last decade and is yet still at an early stage. However, the recent advances in CRP techniques may provide a versatile route to the synthesis of well-defined functional structures and hence facilitate further research in this area. Indeed, responsive properties and functionalities can now be introduced into the nanoreactors without the need of protecting groups or stringent conditions. By tethering the catalysts into a polymeric support the catalyst can be easily recovered but, more importantly, the protected and confined environment provided by the polymeric nanoreactor can act as a yocto-litre reaction vessel concentrating reactants into the core of the confined environment. Hence, solvophobic products can be protected from degradation, the solubility of the catalyst can be modified and the rate of reaction can be enhanced compared with homogeneous systems.
The field of nanoreactors will benefit by further advances in the field of polymer synthesis which in turn will give access to nanoreactors with a higher degree of molecular and compositional homogeneity. Fine control over the sequence and polydispersity of BCPs will open new avenues to mimic the structural complexity and hence the precise and elegant function achieved in natural systems.
However, the synthesis of polymeric nanoreactors must be further simplified to also allow its commercial exploitation and application. Next-generation nanoreactors should be prepared from cheap starting materials in a one pot synthesis enabling for versatile functionalization and result in robust structural characteristics. We see star polymeric nanoreactors as a very promising approach toward this direction.
Acknowledgment
The EPSRC and AWE are thanked for financial support.