
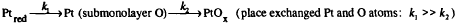

Hostname: page-component-7b7567c8c-wxvwp Total loading time: 0 Render date: 2026-11-17T10:21:06.061Z Has data issue: false hasContentIssue false

Real Time Structural Electrochemistry of Platinum Clusters using Dispersive XAFS

Published online by Cambridge University Press:  15 February 2011

P. G. Allen
Affiliation:
Electronics Research Group, Los Alamos National Laboratory, Los Alamos, NM 87545
S. D. Conradson
Affiliation:
Electronics Research Group, Los Alamos National Laboratory, Los Alamos, NM 87545
M. S. Wilson
Affiliation:
Electronics Research Group, Los Alamos National Laboratory, Los Alamos, NM 87545
S. Gottesfeld
Affiliation:
Electronics Research Group, Los Alamos National Laboratory, Los Alamos, NM 87545
I. D. Raistrick
Affiliation:
Electronics Research Group, Los Alamos National Laboratory, Los Alamos, NM 87545
Get access

Abstract

Chemical reference tables state that the standard potential for the reaction of Pt with water, Pt + 2H2O → Pt(OH)2 + 2H+ + 2e−, is 0.98 V, and electrochemical studies propose that this reaction may occur at potentials as low as 0.8 V [1–5]. Using dispersive x-ray absorption finestructure (XAFS) spectroscopy [6], we have directly probed the structural evolution of a Pt catalyst operating in-situ in a polymer electrolyte [7] fuel cell during cyclic voltammetry. The changes in the number of Pt and O nearest-neighbors and the Pt charge demonstrate a close correspondence with features in the voltammogram. Because dispersive XAFS is very sensitive to detecting structural changes, we have been able to detect the presence of chemisorbed oxygen at potentials of 0.6–0.9 V in the anodic sweep. Since double-layer charging is regarded as the only process in this region for bulk Pt [3–5], these results may reflect a limitation of previous (indirect) studies on Pt electrochemistry, or they may indicate that these clusters are different from their bulk metal counterparts. Exploiting the timeresolving capability of dispersive XAFS, we also monitored changes in the Pt charge and the number of O and Pt nearest-neighbors during the electrochemical oxidation and reduction of the Pt clusters in real-time. The results are inconsistent with those expected from the placeexchange mechanism [8–11] for the formation of the surface oxide on bulk Pt electrodes in aqueous solution;

Our current model for understanding these behaviors is that, relative to bulk Pt, unusual types of surface sites play a major role in determining the reactivity of these clusters.

Access options

Get access to the full version of this content by using one of the access options below. (Log in options will check for institutional or personal access. Content may require purchase if you do not have access.)

Article purchase

Temporarily unavailable