Introduction
Plant wax n-alkanes are commonly used as markers for estimation of intake, digestibility and diet composition in grazing animals (Mayes et al., Reference Mayes, Lamb and Colgrove1986; Dove and Mayes, Reference Dove and Mayes1991; Oliván et al., Reference Oliván, Ferreira, Celaya and Osoro2007), although there are still some concerns to be dealt with. Among them, a major consideration is the need to adjust the faecal alkane concentration for incomplete recovery (Dove and Mayes, Reference Dove and Mayes2005), even for intake estimates (Keli et al., Reference Keli, Andueza, de Vega and Guada2008). Incomplete recovery can be modified by a series of factors including diet type (Dove et al., 1989; Robaina et al., Reference Robaina, Grainger, Moate and Davis1993; Ferreira et al., Reference Ferreira, Oliván, Rodrigues, García and Osoro2005; Charmley and Dove, Reference Charmley and Dove2007) and level of intake (Stakelum and Dillon, 1990; Oliván et al., Reference Oliván, Ferreira, Celaya and Osoro2007), and seems to be due mainly to losses in post-ruminal compartments (Mayes et al., 1988; Kafilzadeh and Parker, Reference Kafilzadeh and Parker1990; Ohajuruka and Palmquist, Reference Ohajuruka and Palmquist1991). Although there is a general agreement about the fact that losses in the ruminal compartment are minimal, if any, Keli (2006) has recently shown that a significant fraction of long-chain n-alkanes is carried to the duodenum linked to, or incorporated into rumen bacteria, but the origin and fate of these alkanes are uncertain. Long-chain n-alkanes have also been used as duodenal flow (Askar et al., Reference Askar, Guada, Balcells, de Vega and Castrillo2005) or rumen transit markers (Giráldez et al., Reference Giráldez, Lamb, López and Mayes2004), but whether the rumen microbes are able to synthesize and/or degrade them in anaerobic conditions remains a question to be answered before these hydrocarbons can be confidently used for these purposes.
McCarthy (Reference McCarthy1964) and Bartley et al. (Reference Bartley, Helmer and Meyer1971) demonstrated that rumen bacteria from sheep and goats are able to incorporate 14C-labelled octadecane (C18) in their lipids. Evidence for bacterial metabolism of the incorporated alkane was discounted because of the absence of radioactivity appearing in volatile fatty acids (VFA), and also because of the anaerobic conditions prevailing in the rumen. Oxygen is needed to form the hydroxyl group required for the oxidation of n-alkanes (Sporman and Widdel, Reference Sporman and Widdel2000). Although many species of bacteria and fungi may oxidize short-chain alkanes (Radwan and Sorkhoh, Reference Radwan and Sorkhoh1993; Sporman and Widdel, Reference Sporman and Widdel2000), none has been isolated in the rumen (Ohajuruka and Palmquist, Reference Ohajuruka and Palmquist1991).
Published papers agree about the dietary origin of tissue n-alkanes in a variety of animal species (Tulliez and Bories, Reference Tulliez and Bories1975a and Reference Tulliez and Bories1975b; Di Muccio et al., Reference Di Muccio, Lintas, Filos and Bernardini1984; Tejeda et al., Reference Tejeda, García, Petrón, Andrés and Antequera2001; Petrón et al., Reference Petrón, Antequera, Muriel, Tejeda and Ventanas2004) although only a few have dealt specifically with this matter in ruminants (Bartley et al., Reference Bartley, Helmer and Meyer1971; Mayes et al., 1988; Ohajuruka and Palmquist, Reference Ohajuruka and Palmquist1991). In rats, radioactive 14C29 can be transformed in long-chain fatty acids, cholesterol esters, di- and tri-glicerides or phospholipids in the liver (Kolattukudy and Hankin, Reference Kolattukudy and Hankin1966). However, the potential for such processes occurring in the ruminants’ foregut has not been explored. It is also possible that n-alkanes of a particular carbon chain length may originate from longer chain n-alkanes.
Literature about these aspects is scarce, if any, despite the fact that they may have an influence on both duodenal and faecal recovery of n-alkanes, and as such on the accuracy of estimates of intake, digestibility, diet composition, duodenal flow or rate of passage obtained using these markers. Hence the objectives of the present experiment were as follows: 1. To establish whether rumen bacteria can synthesize n-alkanes from carbon structures other than waxes (e.g. sugars). 2. To establish whether n-alkanes may be metabolized by rumen bacteria to other components or to shorter chain hydrocarbons. 3. To establish whether rumen bacteria can synthesize n-alkanes from non-alkane wax components (long-chain fatty alcohols, long-chain fatty acids and esters). Owing to the nature of the experiment, the in vitro approach was chosen.
Material and methods
Labelling n-alkanes with 14C
Ryegrass (Lolium perenne) was chosen as the donor of 14C-labelled n-alkanes. For this purpose, the plants were grown in a growth chamber, using a 14/10 h of light/dark cycle, at a constant temperature of 18°C. Turves with actively growing grass were placed inside a large transparent polyethene bag (to contain the radioactive CO2 within a relatively small volume) with the ends connected to a pump, which allowed recirculation of air enriched in 14CO2. The labelled CO2 was obtained by the addition of solid Ba14CO3 (Product no. 297143-1MCI; Sigma-Aldrich, Inc.) to a Petri dish containing 10 ml of frozen (−80°C) 1 M HCl. As the acid thawed, it reacted with the Ba14CO3, releasing 14CO2 into the bag. The labelling period was set to 24 h (20 h of light and 4 h of darkness). After the labelling period, an absorption tube containing a 1 M NaOH solution was connected to the air pump for 3 h with the purpose of capturing any residual 14CO2. The ryegrass was then withdrawn from the polyethene bag and allowed to grow in the growth chamber for at least 1 week. The purpose of this latter step was to ensure the full incorporation of 14C to the plant waxes. Good revisions on the biosynthesis of plant wax n-alkanes can be found in Hawke (Reference Hawke1973), Kolattukudy (Reference Kolattukudy1970), Lester (Reference Lester1979) or Kunst and Samuels (Reference Kunst and Samuels2003), among others. The grass was harvested with scissors and placed in a small polyethene bag for storage at −20°C in the radioisotope laboratory at the Macaulay Institute, prior to freeze-drying and extraction of wax components.
The freeze-dried labelled ryegrass was separated in three fractions:
1. 14C-labelled n-alkanes, which were extracted as follows: the dried plants were extracted with n-heptane, using a Soxhlet apparatus, in sufficient amounts to ensure enough material for the incubations, and without mechanical processing. The extractions were performed until the heptane added was colourless, and at this point full recovery of plant wax components was assumed. The extract, containing all wax components, was applied to a large silica-gel column (200 ml), which had been washed with heptane (3× 150 ml). The n-alkanes were eluted with 3× 150 ml more n-heptane.
2. The solid residue from the Soxhlet extraction, which was considered to be a 14C-labelled plant fraction free of wax components.
3. n-Alkane-free waxes, which were obtained by washing the silica-gel column used in the first extraction with 3× 150 ml of a mixture of n-heptane and ethanol (1 : 1, v/v).
To provide an unlabelled substrate for the in vitro incubations with buffered rumen liquor, turves of perennial ryegrass were grown under similar conditions to those applied to labelled ryegrass (using unlabelled BaCO3), and the same three fractions were obtained.
Experimental procedure
In vitro incubations were carried out in screw-capped 200 mm × 22 mm Pyrex tubes to which 20 ml of a 2 : 1 (v/v) mixture of media solution and inoculum consisting of rumen fluid collected from two fistulated sheep fed on grass hay, transferred into pre-warmed thermos flasks, and strained through four layers of gauze, were added. The media solution was made up with 474 ml distilled water, 0.12 ml microminerals solution (13.2 g CaCl2.2H2O, 10 g MnCl2.4H2O, 1 g CoCl2.6H2O and 0.8 g FeCl2.6H2O in 100 ml distilled water), 237 ml buffer solution (35 g NaHCO3 and 4 g (NH4)HCO3 in 1 l distilled water), 237 ml macrominerals solution (5.7 g Na2HPO4, 6.2 g KH2PO4 and 0.6 g MgSO4.7H2O in 1 l distilled water), 1.22 ml resazurine (0.1 g/100 ml distilled water) and 49 ml reducing solution (285 mg Na2S.7H2O, 2 ml NaOH 1 N and 47.5 ml distilled water). Manipulation was done under continuous flushing of CO2. Tubes were kept warm in a water bath at 38°C and connected to small collapsible plastic (PVC, polyvinyl chloride) bags (those used for serum administration) to collect the CO2 produced during fermentation. Incubation time was set to 48 h. Three different treatments were considered:
1. Incubation of three tubes containing 50 mg each (ca 75 737 640 Bq) of the labelled plant material free of wax components, and 150 mg of non-labelled ryegrass. The aim of this treatment was to test the hypothesis of synthesis of alkanes from organic plant components other than waxes (e.g. sugars).
2. Incubation of three tubes containing 50 mg each of the non-labelled plants free of wax components, plus 0.5 ml of a heptanoic solution containing the 14C-labelled alkanes (ca 1 484 100 Bq) and 150 mg of non-labelled ryegrass. After the addition of the labelled alkanes solution, the heptane was evaporated at room temperature overnight to avoid its possible effect on the incubations. The aim of the treatment was to check the hypothesis of metabolism of alkanes to other components.
3. Incubation of three tubes containing 50 mg each of the non-labelled plants free of wax components, plus 1 ml of a heptanoic solution containing the 14C-labelled n-alkane-free waxes (ca 57 474 720 Bq) and 150 mg of non-labelled ryegrass. After the addition of the labelled alkanes-free waxes solution, the heptane was also evaporated as above. The aim of this treatment was to test the hypothesis of synthesis of alkanes from other wax components.
At the end of the incubation period, 5 ml of 1 M NaOH was added to each serum bag to trap the CO2, allowing a contact time of 24 h. The incubation residues were then treated with 2 ml of H2SO4 1 M to release CO2 from the buffer. After allowing the NaOH in each serum bag to trap the CO2 for 24 h, the bag was sealed and removed from the tube, and 2 ml of 2 M KOH was added to each tube to convert VFA to non-volatile salts. The tubes were frozen at −20°C and then at −80°C prior to freeze-drying. Freeze-dried material was used for n-alkane and VFA analysis.
Sample preparation and analysis
n-Alkanes extraction
Extraction of n-alkanes from residues of reactions was performed in the same Pyrex tubes used for in vitro incubations, using n-docosane (C22) and n-tetratriacontane (C34) as internal standards (0.1 g of a solution containing 0.2 mg of each hydrocarbon per gram. For details, see Mayes et al. (Reference Mayes, Lamb and Colgrove1986) as modified by Dove and Mayes (Reference Dove and Mayes2006)). After evaporation of the filtrate, 2 ml of a solution of n-heptatriacontane (C37) in n-heptane (0.02 mg/g) was added to the bottles as an additional standard in order to quantify any losses of C34 occurring during the extraction process. After redissolving the n-alkanes in the vials (heating at 70°C), 50 μl was transferred to chromatography vials for analysis, adding 10 ml of a pseudocumene-based scintillation fluid suitable for water-insoluble lipid compounds (Insta-fluor Plus; Perkin Elmer Inc., Waltham, MA, USA) to the remainder in preparation for counting for 14C radioactivity.
CO2 analysis
The 5 ml of 1 M NaOH added to serum bags to capture CO2 was transferred to pre-weighed 10 ml test tubes after the set contact time of 24 h. Bags were rinsed with 1 ml of a BaCl2 solution (20% w/v) and 2 ml of a solution of NH4Cl (5% w/v), incorporating the washings to the test tubes. Those were stoppered, shaken and then centrifuged at 2500 r.p.m. (1000 × g) for 10 min. The supernatant was discarded and the precipitate (BaCO3) washed twice with 8 ml of distilled water, repeating the shaking and centrifuging processes. The precipitate was dried by placing unstoppered tubes in an oven at 100°C for 24 h. The tubes were reweighed to determine the amount of BaCO3 obtained, and 20 mg were weighed into scintillation vials to which 2 ml of a 0.2 M solution of sodium-ethylene glycol-bis(beta-aminoethyl ether)-tetraacetic acid (EGTA-Na4) (Sigma-Aldrich, Inc.) was added. The scintillation vials were left (capped) at room temperature for 2 to 3 days, until the BaCO3 was dissolved without the need to shake the vials. Then 12 ml of a scintillator solution for aqueous biological fluids (Hionic-Fluor; Perkin Elmer Inc.) was added to each vial, prior to 14C counting.
VFA extraction
After the extraction of n-alkanes, the residues remaining in the Pyrex tubes were transferred to Erlenmeyer flasks, and 10 ml of 10% H2SO4 added to ensure acidity. The flasks were frozen at −80°C and attached to glass lyophiliser tubes, which were immersed in liquid nitrogen. By attaching a vacuum pump to the lyophiliser tubes, the contents of the Erlenmeyer flasks were freeze-dried, with the water and other volatiles (including VFA) being collected in the lyophiliser tubes. When samples were freeze-dried, the pump was switched off and the ends of the lyophiliser tubes were covered with a plastic film until they reached room temperature. Then contents of the tube (containing VFA) were transferred to scintillation vials to which 2 ml of 1 M NaOH was added. The vials were dried at 70°C and the residue dissolved in 2 ml of distilled water. Hionic-Fluor scintillation fluid (12 ml) was then added prior to 14C counting.
Measurement of 14C radioactivity
For determination of the amount of 14C in whole and de-waxed labelled dried grass, samples were prepared for a liquid scintillation counter (Permafluor) using a simple oxidizer (Model 306 Packard Oxidizer) available at the Macaulay Institute.
The 14C radioactivity in labelled samples (alkanes, CO2 and VFA) and whole and de-waxed labelled dried grass was determined using a Liquid Scintillation Analyser (Tri-Carb 2900TR; Perkin Elmer Inc.). The count rates were corrected for quenching using the external standard method with the tSIE (transformed spectral index of the external standard) as the quench indication parameter provided by the scintillation counter, and employing separate tSIE/efficiency regressions for each type of scintillation fluid.
Results
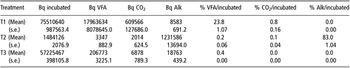
Table 1 shows the quantities of 14C radioactivity (Bq) in the materials used in the three incubation treatments described in the experimental procedure, and the partitioning of this activity between the different samples obtained after 48 h of incubation. In Treatment 1 (incubation of labelled plants free of wax components), the greater activity was found in VFA (23.8% of that of the incubated material), whereas only 0.8% was found in CO2 and none in n-alkanes. In Treatment 2 (incubation of labelled alkanes), the greater activity corresponded to n-alkanes (83.0%) and was very low in VFA (0.2%) and CO2 (0.1%). In the third treatment (incubation of labelled n-alkane-free waxes), negligible 14C activity was detected in n-alkanes (0.0%), CO2 (0.0%) or VFA (0.4%).
14C activity in the incubated material used in the three experimental treatments (T1, T2 and T3) and in volatile fatty acids (VFA), CO2 and n-alkanes (Alk) obtained after in vitro incubation for 48 h (n = 3)

Bq = becquerel; T1 = incubation of labelled ryegrass free of wax components; T2 = incubation of labelled n-alkanes; T3 = incubation of labelled n-alkane-free waxes.
Discussion
In order to use long-chain n-alkanes confidently as either duodenal flow or rumen transit markers, losses in the foregut must be investigated, including the possible role of rumen bacteria. In this context, the objective of the present experiment was to test whether within a rumen liquor fermentation system n-alkanes can be derived from the de-waxed structures of the plant or from non-alkane wax components (long-chain fatty alcohols, long-chain fatty acids and esters), or may be metabolized by bacteria to other components or to shorter-chain hydrocarbons. Ohajuruka and Palmquist (Reference Ohajuruka and Palmquist1991) observed that almost 15% of some of the ingested n-alkanes disappeared from the forestomachs under controlled feeding conditions in dairy cows.
In Treatment 1, respective recoveries of 14C activity in VFA and CO2 were 68% and 16% of the theoretical values. These latter were calculated assuming an efficiency of the microbial synthesis of 21.5 g N/kg fermentable organic matter, and a bacterial protein content of 51% in the dry matter (INRA, 1978). It can also be estimated that 74% of the degraded substrate is used for fermentation, and the rest is incorporated by the growing bacterial population. Taking into account the stoichiometry of fermentation (Hungate Reference Hungate1966), and the molecular weight of fermentation products, it can be calculated that VFA should account for 68%, CO2 for 16% and methane for 5% of the fermentation products. Then, for a 60% degradability VFA should represent, approximately, 30% of the incubated material, and CO2 about 7%. Bearing in mind that 14C data will only refer to carbon, and assuming that forage dry matter (as cellulose) contains 40% C, CO2 contains 27.3% C and VFA about 47% C, 35% of the incubated C should be recovered in VFA, and nearly 5% in CO2. The difference between these values and those shown in Table 1 (Treatment 1) can be attributed, at least in part, to possible physical losses of the incubated material through the process of incubation and sample manipulation, as the amount of labelled material used was very small. Moreover, this difference between calculated and recorded radioactivity recovery in CO2 cannot be explained only by these likely losses. An incomplete dissolution of BaCO3 in EGTA-Na4, or possible CO2 losses from serum bags, might also have occurred. Another possible reason for low 14C values could be that the fermentation was not complete, as the ordinary gas production method carries on for about 96 h.
Results from the first and third treatments show that there was no microbial synthesis of n-alkanes from other wax components (alcohols, long-chain fatty acids and esters in Treatment 3) or carbon structures other than waxes (e.g. carbohydrates; Treatment 1). This is in contrast with the synthesis process of n-alkanes in the plant, where C16 and C18 long-chain fatty acids are formed de novo from acetate in the chloroplasts (Lester, Reference Lester1979), further following elongation and either decarboxylation or condensation of at least two fatty acids (Hawke, Reference Hawke1973). Results from Treatment 2 show that there was no n-alkane degradation to CO2 or VFA, although the 14C activity was not fully recovered (83%). The source of error could have been likely losses of n-alkanes through the extraction process. However, this explanation was discarded, as the comparison between the concentrations of C34 (added before the saponification) and C37 (added after extraction) did not support possible alkane losses (average [(weight of C37 before incubation/concentration of C37 after incubation)/(weight of C34 before incubation/concentration of C34 after incubation)] = 1.01). As in Treatment 1, the possible physical losses of the incubated material through the process of incubation and sample manipulation can be blamed, at least in part.
There seems to be a lack of information about the metabolism of n-alkanes in in vitro studies in the literature, and hence our results were compared with those obtained in vivo. To this respect, our results strongly suggested that ruminal bacteria are not able to synthesize n-alkanes from other wax components, and that they are not metabolized by the ruminal microbiota. The same conclusion was reached in vivo by McCarthy (Reference McCarthy1964) and Bartley et al. (Reference Bartley, Helmer and Meyer1971), working with 14C-octadecane in goats and cattle, respectively, although they found high concentrations of the hydrocarbon in isolated bacterial extracts. Their hypothesis was that ruminal bacteria may incorporate alkanes in their lipids, maybe by passive diffusion, but without further metabolism. However, McCarthy (Reference McCarthy1964) demonstrated the conversion of the octadecane to the corresponding fatty acid during its absorption through the ruminal wall in perfusion studies. A similar process is still to be demonstrated with alkanes of longer chain or in sheep or cattle, where in vivo results show that ruminal disappearance of alkanes is small, if any (Mayes et al., 1988; Ohajuruka and Palmquist, Reference Ohajuruka and Palmquist1991). To this respect, Mayes et al. (1988) indicated that C32 was oxidized in sheep but they did not give any information about the place or the mechanism, proposing a similar process to that described for C29 in rats by Kolattukudy and Hankin (Reference Kolattukudy and Hankin1966), who demonstrated its transformation in the liver into n-17 lipids, cholesterol esters, di- and tri-glicerides and phospholipids.
Conclusions
The results from the present experiment demonstrated that there is neither synthesis nor degradation of n-alkanes by ruminal bacteria in vitro. This supports the results obtained in vivo in previous papers, in which significant ruminal disappearance of n-alkanes was discarded. Hence, the long-chain n-alkanes can be confidently used as duodenal flow or rumen transit markers regardless of their incomplete faecal recovery.
Acknowledgements
Mr. A. Keli was awarded a fellowship of the ‘Plan de Formación de Personal Investigador (FPI)’ of the Spanish Ministry for Education and Science (project no. AGL2001-2771) to carry out his stay at the Macaulay Institute. Thanks are also due to A. Sim and E. Green at the Macaulay Institute, who provided practical advice and assistance.