1 Introduction
The use of electric fields to drive flow parallel to charged surfaces at the micro- and nano-scale is ubiquitous in science and technology, with applications in numerous fields, from ‘Lab-on-a-Chip’ devices (Stone, Stroock & Ajdari Reference Stone, Stroock and Ajdari2004; Squires & Quake Reference Squires and Quake2005) and nano-fluidics (Bocquet & Charlaix Reference Bocquet and Charlaix2010) to membrane and soil science, and the separation and analysis of biological macromolecules (Eijkel & Berg Reference Eijkel and van den Berg2005). In its simplest form, the fluid velocity
 $v_{\infty }$
far from the charged surface is proportional to the applied electric field
 $E_{0}$
,
 $v_{\infty }=\unicode[STIX]{x1D707}_{e}E_{0}$
. The electro-osmotic mobility
 $\unicode[STIX]{x1D707}_{e}$
is given by the Helmholtz–Smoluchowski equation (von Smoluchowski Reference von Smoluchowski1903),
 $\unicode[STIX]{x1D707}_{e}=-\unicode[STIX]{x1D700}\unicode[STIX]{x1D701}/\unicode[STIX]{x1D702}$
, where
 $\unicode[STIX]{x1D700}$
is the solvent permittivity,
 $\unicode[STIX]{x1D702}$
is the solvent viscosity, and the so-called zeta potential
 $\unicode[STIX]{x1D701}$
is the electric potential at the shear plane. In neat (single-component) solvents, this electro-osmotic flow (EOF) has been studied extensively (Delgado et al.
Reference Delgado, González-Caballero, Hunter, Koopal and Lyklema2007), with more recent works focusing on the influence of the channel geometry (Bhattacharyya, Zheng & Conlisk Reference Bhattacharyya, Zheng and Conlisk2005; Mao, Sherwood & Ghosal Reference Mao, Sherwood and Ghosal2014), surface slip (Bouzigues, Tabeling & Bocquet Reference Bouzigues, Tabeling and Bocquet2008; Huang et al.
Reference Huang, Cottin-Bizonne, Ybert and Bocquet2008; Maduar et al.
Reference Maduar, Belyaev, Lobaskin and Vinogradova2015; Rankin & Huang Reference Rankin and Huang2016), ion specificity (Huang et al.
Reference Huang, Cottin-Bizonne, Ybert and Bocquet2007) and solvent rheology (Bautista et al.
Reference Bautista, Sánchez, Arcos and Méndez2013). Electro-osmotic flow in solvent mixtures is common in non-aqueous capillary electrophoresis (Kenndler Reference Kenndler2014), and has been investigated also for aqueous mixtures (Valkó, Sirén & Riekkola Reference Valkó, Sirén and Riekkola1999; Grob & Steiner Reference Grob and Steiner2002), but the electrokinetics in such systems is not well understood.
The difference between the electrostatics of neat solvents and solvent mixtures has also been an area of intense research in recent years, with the growing use of miscible and non-miscible oil–water mixtures in colloidal science and microfluidics. A crucial role is played by the partitioning of ions between solvents, due to their preferential solvation in one of the liquids (Onuki & Kitamura Reference Onuki and Kitamura2004; Tsori & Leibler Reference Tsori and Leibler2007; Zwanikken & van Roij Reference Zwanikken and van Roij2007; Araki & Onuki Reference Araki and Onuki2009; Ben-Yaakov et al.
Reference Ben-Yaakov, Andelman, Harries and Podgornik2009; Bier et al.
Reference Bier, Gambassi, Oettel and Dietrich2011; Okamoto & Onuki Reference Okamoto and Onuki2011; Pousaneh & Ciach Reference Pousaneh and Ciach2011; Samin & Tsori Reference Samin and Tsori2011, Reference Samin and Tsori2013; Bier, Gambassi & Dietrich Reference Bier, Gambassi and Dietrich2012; Samin & Tsori Reference Samin and Tsori2012; Pousaneh & Ciach Reference Pousaneh and Ciach2014; Michler et al.
Reference Michler, Shahidzadeh, Westbroek, van Roij and Bonn2015; Samin & Tsori Reference Samin and Tsori2016). The preferential wetting of one liquid at a solid surface or the presence of a liquid–liquid interface therefore also affects the electrostatics of the mixture. It leads to a modification of colloid–colloid (Law, Petit & Beysens Reference Law, Petit and Beysens1998; Hertlein et al.
Reference Hertlein, Helden, Gambassi, Dietrich and Bechinger2008; Bonn et al.
Reference Bonn, Otwinowski, Sacanna, Guo, Wegdam and Schall2009; Nellen et al.
Reference Nellen, Dietrich, Helden, Chodankar, Nygård, van der Veen and Bechinger2011; Samin et al.
Reference Samin, Hod, Melamed, Gottlieb and Tsori2014) as well as colloid–interface interactions (Leunissen et al.
Reference Leunissen, van Blaaderen, Hollingsworth, Sullivan and Chaikin2007a
,Reference Leunissen, Zwanikken, van Roij, Chaikin and van Blaaderen
b
; Banerjee et al.
Reference Banerjee, Williams, Azevedo, Helgeson and Squires2016; Elbers et al.
Reference Elbers, van der Hoeven, de Winter, Schneijdenberg, van der Linden, Filion and van Blaaderen2016; Everts, Samin & van Roij Reference Everts, Samin and van Roij2016). The strength of preferential solvation is measured by the Gibbs transfer energy
 $k_{B}Tg^{\unicode[STIX]{x1D6FC}}$
of an ion species
 $\unicode[STIX]{x1D6FC}$
between two solvents, where
 $k_{B}T$
is the thermal energy and
 $|g^{\unicode[STIX]{x1D6FC}}|\sim 1{-}10$
for aqueous mixtures of relatively polar organic solvents (Kalidas, Hefter & Marcus Reference Kalidas, Hefter and Marcus2000; Marcus Reference Marcus2007), but can be as large as 15 in less polar solvents containing antagonistic salts (Onuki et al.
Reference Onuki, Yabunaka, Araki and Okamoto2016). For monovalent salts dissolved in mixtures, the combined effect of both ionic species (Bier et al.
Reference Bier, Gambassi, Oettel and Dietrich2011; Okamoto & Onuki Reference Okamoto and Onuki2011; Samin & Tsori Reference Samin and Tsori2013; Michler et al.
Reference Michler, Shahidzadeh, Westbroek, van Roij and Bonn2015) is conveniently expressed in terms of the average overall solubility
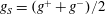 $g_{S}=(g^{+}+g^{-})/2$
and the solubility contrast
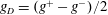 $g_{D}=(g^{+}-g^{-})/2$
, which, for example, determines the Donnan potential
 $k_{B}Tg_{D}/e$
at oil–water interfaces due to ion partitioning, where
 $e$
is the elementary charge. In this work, we show that preferential solvation also affects electrokinetic phenomena in mixtures, where it generates an additional, and significant, source of fluid mobility. We find that the dominant contribution to this additional mobility is proportional to
 $g_{D}$
and independent of the surface charge, and is therefore able to generate flow even at electrically neutral surfaces.
2 Formulation of the problem
Within a continuum theory, we study the flow in a channel containing an electrolytic mixture using direct numerical simulations and a simple linear theory. We consider an oil–water mixture characterized by the order parameter
 $\unicode[STIX]{x1D711}$
, which is the deviation of the volume fraction of water in the mixture,
 $\unicode[STIX]{x1D719}$
, from its critical value
 $\unicode[STIX]{x1D719}_{c}$
:
 $\unicode[STIX]{x1D711}=\unicode[STIX]{x1D719}-\unicode[STIX]{x1D719}_{c}$
. The mixture contains point-like monovalent ions with number densities
 $n^{\pm }$
. In our calculations, two cylindrical reservoirs containing the mixture are coaxially connected by a long cylindrical channel with radius
 $R$
, with the
 $z$
axis being the axis of symmetry and
 $r$
the radial coordinate. In equilibrium, the composition in both of the reservoirs is
 $\unicode[STIX]{x1D711}_{0}=0$
and the number densities of ions are
 $n_{0}$
. At the edge of one reservoir, we impose a uniform external electric field
 $E_{0}\hat{z}$
, forcing the charged mixture in the channel, and leading to a flow field
 $\boldsymbol{v}$
.
2.1 Governing equations
To study the mixture dynamics, we start from a free energy
 $F$
of the form
 $$\begin{eqnarray}\displaystyle \unicode[STIX]{x1D6FD}F & = & \displaystyle \int \text{d}\boldsymbol{r}\Bigg\{f_{m}(\unicode[STIX]{x1D711})+\frac{C}{2}|\unicode[STIX]{x1D735}\unicode[STIX]{x1D711}|^{2}+\frac{\unicode[STIX]{x1D6FD}}{2}\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})(\unicode[STIX]{x1D735}\unicode[STIX]{x1D713})^{2}\nonumber\\ \displaystyle & & \displaystyle +\mathop{\sum }_{\unicode[STIX]{x1D6FC}=\pm }n^{\unicode[STIX]{x1D6FC}}[(\log (v_{0}n^{\unicode[STIX]{x1D6FC}})-1)-g^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D711}]\Bigg\},\end{eqnarray}$$
where
 $\unicode[STIX]{x1D6FD}^{-1}=k_{B}T$
and
 $v_{0}=a^{3}$
is the molecular volume of both mixture components. The first term in the integrand is the ‘double-well’ bulk mixture free energy density,
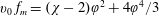 $v_{0}\,f_{m}=(\unicode[STIX]{x1D712}-2)\unicode[STIX]{x1D711}^{2}+4\unicode[STIX]{x1D711}^{4}/3$
, where
 $\unicode[STIX]{x1D712}\sim 1/T$
is the Flory parameter. This free energy leads to an upper critical solution temperature type phase diagram, with a critical temperature
 $T_{c}$
and the corresponding critical Flory parameter
 $\unicode[STIX]{x1D712}_{c}=2$
. In this work, we focus on the region
 $T>T_{c}$
(
 $\unicode[STIX]{x1D712}<\unicode[STIX]{x1D712}_{c}$
), such that the bulk mixture is always homogeneous. The second ‘square gradient’ term accounts for composition inhomogeneities at interfaces, where
 $C=\unicode[STIX]{x1D712}/a$
. The third term is the electrostatic energy density, where
 $\unicode[STIX]{x1D713}$
is the electric potential and
 $\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})$
is the permittivity, assumed to depend linearly on
 $\unicode[STIX]{x1D711}$
. The final term in the integrand is the ionic free energy, composed of the ideal-gas entropy of ions of species
 $\unicode[STIX]{x1D6FC}=\pm$
and the ionic solvation energy, which is proportional to the Gibbs free energy of transfer
 $g^{\unicode[STIX]{x1D6FC}}$
and local solvent composition. It should be noted that
 $g^{\unicode[STIX]{x1D6FC}}$
can vary greatly between ionic species depending on their size, charge and chemistry.
The relations governing the mixture dynamics read (Araki & Onuki Reference Araki and Onuki2009)
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x2202}\unicode[STIX]{x1D711}/\unicode[STIX]{x2202}t+\unicode[STIX]{x1D735}\boldsymbol{\cdot }(\unicode[STIX]{x1D711}\boldsymbol{v})=D^{m}\unicode[STIX]{x1D6FB}^{2}\unicode[STIX]{x1D6FD}\unicode[STIX]{x1D707}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x2202}n^{\unicode[STIX]{x1D6FC}}/\unicode[STIX]{x2202}t+\unicode[STIX]{x1D735}\boldsymbol{\cdot }(n^{\unicode[STIX]{x1D6FC}}\boldsymbol{v})=D^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D735}\boldsymbol{\cdot }n^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D735}\unicode[STIX]{x1D6FD}\unicode[STIX]{x1D706}^{\unicode[STIX]{x1D6FC}}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D735}\boldsymbol{\cdot }(\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})\unicode[STIX]{x1D735}\unicode[STIX]{x1D713})=-\mathop{\sum }_{\unicode[STIX]{x1D6FC}}z^{\unicode[STIX]{x1D6FC}}n^{\unicode[STIX]{x1D6FC}}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D735}\boldsymbol{\cdot }\boldsymbol{v}=0, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D702}\unicode[STIX]{x1D6FB}^{2}\boldsymbol{v}-\unicode[STIX]{x1D735}p=v_{0}^{-1}\unicode[STIX]{x1D711}\unicode[STIX]{x1D735}\unicode[STIX]{x1D707}+\mathop{\sum }_{\unicode[STIX]{x1D6FC}}n^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D735}\unicode[STIX]{x1D706}^{\unicode[STIX]{x1D6FC}}. & \displaystyle\end{eqnarray}$$
Equation (2.2) is the convective Cahn–Hilliard equation, where
 $D^{m}$
is the interdiffusion constant of the mixture and
 $\unicode[STIX]{x1D707}=v_{0}(\unicode[STIX]{x1D6FF}F/\unicode[STIX]{x1D6FF}\unicode[STIX]{x1D711})$
is the solvent chemical potential, given by
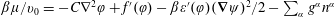 $\unicode[STIX]{x1D6FD}\unicode[STIX]{x1D707}/v_{0}=-C\unicode[STIX]{x1D6FB}^{2}\unicode[STIX]{x1D711}+f^{\prime }(\unicode[STIX]{x1D711})-\unicode[STIX]{x1D6FD}\unicode[STIX]{x1D700}^{\prime }(\unicode[STIX]{x1D711})(\unicode[STIX]{x1D735}\unicode[STIX]{x1D713})^{2}/2-\sum _{\unicode[STIX]{x1D6FC}}g^{\unicode[STIX]{x1D6FC}}n^{\unicode[STIX]{x1D6FC}}$
. Equations (2.3) and (2.4) are the Poisson–Nernst–Planck equations, where the
 $D^{\unicode[STIX]{x1D6FC}}$
are ionic diffusion constants and
 $\unicode[STIX]{x1D706}^{\unicode[STIX]{x1D6FC}}=\unicode[STIX]{x1D6FF}F/\unicode[STIX]{x1D6FF}n^{\unicode[STIX]{x1D6FC}}$
are the ionic chemical potentials given by
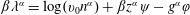 $\unicode[STIX]{x1D6FD}\unicode[STIX]{x1D706}^{\unicode[STIX]{x1D6FC}}=\log (v_{0}n^{\unicode[STIX]{x1D6FC}})+\unicode[STIX]{x1D6FD}z^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D713}-g^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D711}$
, with
 $z^{\unicode[STIX]{x1D6FC}}=\pm e$
. Equations (2.5) and (2.6) are the Stokes equations at small Reynolds number, where
 $p$
is the pressure and the right-hand side of (2.6) contains the body forces due to concentration gradients.
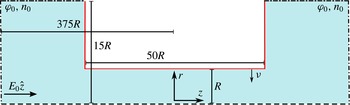
An illustration of the system in the
 $rOz$
plane. A cylindrical channel with radius
 $R$
connects two reservoirs containing a salty water–acetonitrile mixture with a composition
 $\unicode[STIX]{x1D711}_{0}=0$
and a salt concentration
 $n_{0}$
. The system is driven out of equilibrium by an external electric field
 $E_{0}$
in the
 $\hat{z}$
direction. The system dimensions, given in the figure in terms of
 $R$
, ensure that the effect of reservoirs and channel edges on the flow that develops in the channel is small. Different boundary line types indicate different boundary conditions (see text).

An illustration of the computational domain for the numerical solution of (2.2)–(2.6) is shown in figure 1 (not to scale). Solid walls are indicated by the red lines in figure 1. On these walls, we impose the no-slip boundary condition (BC) for the velocity,
 $\boldsymbol{v}=0$
, and no material fluxes for the composition and ions. The second BC for the composition is
 $-\unicode[STIX]{x1D742}\boldsymbol{\cdot }\unicode[STIX]{x1D735}\unicode[STIX]{x1D711}=\unicode[STIX]{x1D6FE}$
, with
 $\unicode[STIX]{x1D742}$
being the unit normal to the surface. The coefficient
 $\unicode[STIX]{x1D6FE}$
represents the short-range interaction between the solvent mixture and the surface (per solvent molecule). We call
 $\unicode[STIX]{x1D6FE}$
the effective surface field;
 $\unicode[STIX]{x1D6FE}$
is positive (negative) for hydrophilic (hydrophobic) surfaces. The surface may carry a fixed charge density
 $e\unicode[STIX]{x1D70E}$
, which from Gauss’s law implies
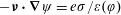 $-\unicode[STIX]{x1D742}\boldsymbol{\cdot }\unicode[STIX]{x1D735}\unicode[STIX]{x1D713}=e\unicode[STIX]{x1D70E}/\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})$
.
At the open reservoir edges, indicated by dash–dot lines in figure 1, we set the composition to
 $\unicode[STIX]{x1D711}_{0}=0$
and the ion densities to
 $n_{0}$
. We also allow the mixture to be freely advected, with vanishing total stress and diffusive fluxes,
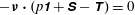 $-\unicode[STIX]{x1D742}\boldsymbol{\cdot }(p\unicode[STIX]{x1D7ED}+\unicode[STIX]{x1D64E}-\unicode[STIX]{x1D64F})=0$
,
 $-\unicode[STIX]{x1D742}\boldsymbol{\cdot }\unicode[STIX]{x1D735}\unicode[STIX]{x1D707}=0$
and
 $-\unicode[STIX]{x1D742}\boldsymbol{\cdot }n^{\pm }\unicode[STIX]{x1D735}\unicode[STIX]{x1D706}^{\pm }=0$
respectively, where
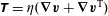 $\unicode[STIX]{x1D64F}=\unicode[STIX]{x1D702}(\unicode[STIX]{x1D735}\boldsymbol{v}+\unicode[STIX]{x1D735}\boldsymbol{v}^{\text{T}})$
is the viscous stress tensor and
 $\unicode[STIX]{x1D64E}$
is the total stress tensor, given by
 $$\begin{eqnarray}\displaystyle \unicode[STIX]{x1D6FD}\unicode[STIX]{x1D64E} & = & \displaystyle \left[\unicode[STIX]{x1D711}f_{m}^{\prime }(\unicode[STIX]{x1D711})-f_{m}-\frac{C}{2}|\unicode[STIX]{x1D735}\unicode[STIX]{x1D711}|^{2}-C\unicode[STIX]{x1D711}\unicode[STIX]{x1D6FB}^{2}\unicode[STIX]{x1D711}-\unicode[STIX]{x1D711}\unicode[STIX]{x1D700}^{\prime }(\unicode[STIX]{x1D711})\frac{|\boldsymbol{E}|^{2}}{2}+\frac{\boldsymbol{E}\boldsymbol{\cdot }\boldsymbol{D}}{2}\right.\nonumber\\ \displaystyle & & \displaystyle \left.+\mathop{\sum }_{\unicode[STIX]{x1D6FC}=\pm }(1-g^{\unicode[STIX]{x1D6FC}}\unicode[STIX]{x1D711})n^{\unicode[STIX]{x1D6FC}}\right]\unicode[STIX]{x1D7ED}+C\unicode[STIX]{x1D735}\unicode[STIX]{x1D711}\unicode[STIX]{x1D735}\unicode[STIX]{x1D711}-\boldsymbol{E}\boldsymbol{D},\end{eqnarray}$$
where
 $\boldsymbol{E}=-\unicode[STIX]{x1D735}\unicode[STIX]{x1D713}$
is the electric field,
 $\boldsymbol{D}=\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})\boldsymbol{E}$
is the displacement field and the prime indicts differentiation with respect to the argument. Lastly, the dashed curve in figure 1 is the
 $r=0$
axis, where we apply symmetry BCs for all fields. The numerical simulations in this work were performed using the finite-elements software COMSOL multiphysics.
2.2 Linear theory
To better understand the flow generated in the channel, we first consider a simplified system where we assume that (i) perturbations in the composition
 $\unicode[STIX]{x1D711}-\unicode[STIX]{x1D711}_{0}$
and ion densities
 $\unicode[STIX]{x1D6FF}n^{\pm }=n^{\pm }-n_{0}$
relative to their bulk values are small due to weak adsorption at the channel surface, (ii) the channel is very long such that edge effects are negligible and translational invariance in the
 $z$
direction applies, (iii) the dominant body force in (2.6) is the electric body force and (iv) convective composition and ion currents in the channel are negligible since the corresponding Péclet numbers are small. However, even in simple mixtures, transport through channels can lead to complex and surprising effects (Samin & van Roij Reference Samin and van Roij2017), also for small Péclet numbers. Therefore, it is necessary to also solve the complete system equations (2.2)–(2.6) numerically to verify that the simplified description is valid.
Within the appropriate parameter regime, the full transport problem can thus be reduced to a steady-state problem, where all fields depend only on
 $r$
. We also decompose the electric potential as
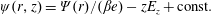 $\unicode[STIX]{x1D713}(r,z)=\unicode[STIX]{x1D6F9}(r)/(\unicode[STIX]{x1D6FD}e)-zE_{z}+\text{const.}$
, where
 $\unicode[STIX]{x1D6F9}(r)$
is a dimensionless radial potential and
 $E_{z}$
is the uniform axial electric field in the channel. It should be noted that
 $E_{z}\neq E_{0}$
in general. For a fully developed flow, we assume that
 $v_{r}=0$
and thus
 $v_{z}=v_{z}(r)$
from (2.5). Lastly, we set
 $\unicode[STIX]{x1D735}p=0$
as we focus on the effect of
 $E_{z}$
. Linearizing the simplified equations (2.2)–(2.6) (Okamoto & Onuki Reference Okamoto and Onuki2011; Samin & Tsori Reference Samin and Tsori2013), we find
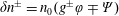 $\unicode[STIX]{x1D6FF}n^{\pm }=n_{0}(g^{\pm }\unicode[STIX]{x1D711}\mp \unicode[STIX]{x1D6F9})$
and obtain the system
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D709}^{2}\unicode[STIX]{x1D6FB}_{r}^{2}\unicode[STIX]{x1D711}=\unicode[STIX]{x1D711}+2n_{0}g_{D}(\unicode[STIX]{x1D6F9}-g_{D}\unicode[STIX]{x1D711})/\unicode[STIX]{x1D70F}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D706}_{D}^{2}\unicode[STIX]{x1D6FB}_{r}^{2}\unicode[STIX]{x1D6F9}=\unicode[STIX]{x1D6F9}-g_{D}\unicode[STIX]{x1D711}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D706}_{D}^{2}\unicode[STIX]{x1D702}\unicode[STIX]{x1D6FB}_{r}^{2}v_{z}=\bar{\unicode[STIX]{x1D700}}(\unicode[STIX]{x1D6F9}-g_{D}\unicode[STIX]{x1D711})E_{z}/(\unicode[STIX]{x1D6FD}e), & \displaystyle\end{eqnarray}$$
where
 $\unicode[STIX]{x1D709}=(C/\unicode[STIX]{x1D70F})^{1/2}$
is the modified correlation length and
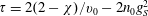 $\unicode[STIX]{x1D70F}=2(2-\unicode[STIX]{x1D712})/v_{0}-2n_{0}g_{S}^{2}$
. The Debye length is
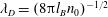 $\unicode[STIX]{x1D706}_{D}=(8\unicode[STIX]{x03C0}l_{B}n_{0})^{-1/2}$
, where
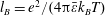 $l_{B}=e^{2}/(4\unicode[STIX]{x03C0}\bar{\unicode[STIX]{x1D700}}k_{B}T)$
is the Bjerrum length with the average permittivity
 $\bar{\unicode[STIX]{x1D700}}=\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711}_{0})$
.
3 Results
In a cylindrical domain, the profiles
 $\unicode[STIX]{x1D711}(r)$
and
 $\unicode[STIX]{x1D6F9}(r)$
of (2.8) and (2.9) are the linear combinations
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D6F9}(r)=a_{1}\text{I}_{0}(q_{1}r)-a_{2}\text{I}_{0}(q_{2}r), & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle \unicode[STIX]{x1D711}(r)=b_{1}\text{I}_{0}(q_{1}r)-b_{2}\text{I}_{0}(q_{2}r), & \displaystyle\end{eqnarray}$$
where
 $\text{I}_{n}$
is the modified Bessel function of the first kind of order
 $n$
, such that the wavenumbers
 $q_{i}$
(
 $i=1,2$
) obey the biquadratic equation
 $$\begin{eqnarray}\unicode[STIX]{x1D706}_{D}^{2}q_{i}^{4}-[1+(\unicode[STIX]{x1D706}_{D}/\unicode[STIX]{x1D709})^{2}-g_{D}^{2}/(4\unicode[STIX]{x03C0}l_{B}C)]q_{i}^{2}+\unicode[STIX]{x1D709}^{-2}=0.\end{eqnarray}$$
The amplitudes
 $a_{i}$
and
 $b_{i}$
follow from symmetry at
 $r=0$
and the BCs given above at the channel surface
 $r=R$
. Also making use of the identity
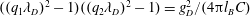 $((q_{1}\unicode[STIX]{x1D706}_{D})^{2}-1)((q_{2}\unicode[STIX]{x1D706}_{D})^{2}-1)=g_{D}^{2}/(4\unicode[STIX]{x03C0}l_{B}C)$
, which follows from (3.3), the amplitudes read
 $$\begin{eqnarray}\displaystyle & \displaystyle a_{i}=\frac{g_{D}}{1-(\unicode[STIX]{x1D706}_{D}q_{i})^{2}}b_{i}, & \displaystyle\end{eqnarray}$$
 $$\begin{eqnarray}\displaystyle & \displaystyle b_{i}=\frac{\unicode[STIX]{x1D6FE}[1-(\unicode[STIX]{x1D706}_{D}q_{i})^{2}]-g_{D}\unicode[STIX]{x1D70E}/C}{\unicode[STIX]{x1D706}_{D}^{2}q_{i}(q_{2}^{2}-q_{1}^{2})\text{I}_{1}(q_{i}R)}. & \displaystyle\end{eqnarray}$$
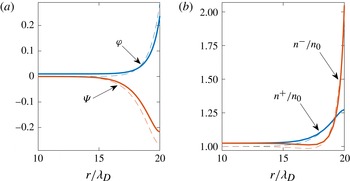
We plot in figure 2(a) the resulting composition and potential profiles for a water–acetonitrile mixture with
 $n_{0}=50$
mM of NaCl (
 $\unicode[STIX]{x1D706}_{D}=1.16$
nm,
 $g^{+}\approx 0$
and
 $g^{-}\approx 4$
(Kalidas et al.
Reference Kalidas, Hefter and Marcus2000; Marcus Reference Marcus2007)) at room temperature (
 $\unicode[STIX]{x1D712}=1.86$
,
 $\unicode[STIX]{x1D709}=1.31$
nm) inside a channel with radius
 $R=20\unicode[STIX]{x1D706}_{D}$
. The mixture physical properties are given in the caption of figure 2. We assume that these properties are independent of temperature, except for the mixture interdiffusion constant
 $D^{m}$
, which follows from
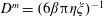 $D^{m}=(6\unicode[STIX]{x1D6FD}\unicode[STIX]{x03C0}\unicode[STIX]{x1D702}\unicode[STIX]{x1D709})^{-1}$
(Kawasaki Reference Kawasaki1970). The ionic diffusion constants are taken to be
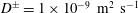 $D^{\pm }=1\times 10^{-9}~\text{m}^{2}~\text{s}^{-1}$
.
(a) Composition
 $\unicode[STIX]{x1D711}(r)$
and scaled potential
 $\unicode[STIX]{x1D6F9}(r)$
at the channel centre (
 $z=0$
) and (b) the corresponding scaled cation and anion profiles, for a water–acetonitrile mixture with a Debye length
 $\unicode[STIX]{x1D706}_{D}=1.16$
nm and a correlation length
 $\unicode[STIX]{x1D709}=1.31$
nm. The solid lines represent the numerical solution and the dashed curves represent the linear theory (see text). We plot the profiles near the channel surface, where here and in all other figures the channel radius is
 $R=20\unicode[STIX]{x1D706}_{D}$
. The channel surface is uncharged (
 $\unicode[STIX]{x1D70E}=0$
) and hydrophilic,
 $\unicode[STIX]{x1D6FE}=0.2C$
, corresponding to a tension of
 ${\approx}7~\text{mN}~\text{m}^{-1}$
. The mixture with
 $\bar{\unicode[STIX]{x1D700}}=58$
and at a temperature of 293 K (
 $\unicode[STIX]{x1D712}=1.86$
) contains 50 mM of NaCl (
 $g^{+}=0,g^{-}=4$
). For the mixture properties, we use the critical temperature
 $T_{c}=272$
K and a molecular length
 $a=3.6$
Å, estimated from a critical density of
 $900~\text{kg}~\text{m}^{-3}$
and a viscosity of 0.7 mPa s (Sazonov et al.
Reference Sazonov, Shaw, Skrzecz, Lisov and Sazonov2007; Wohlfarth Reference Wohlfarth2009). The liquid permeability is given by
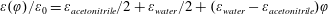 $\unicode[STIX]{x1D700}(\unicode[STIX]{x1D711})/\unicode[STIX]{x1D700}_{0}=\unicode[STIX]{x1D700}_{acetonitrile}/2+\unicode[STIX]{x1D700}_{water}/2+(\unicode[STIX]{x1D700}_{water}-\unicode[STIX]{x1D700}_{acetonitrile})\unicode[STIX]{x1D711}$
, where
 $\unicode[STIX]{x1D700}_{0}$
is the vacuum permittivity, and we used
 $\unicode[STIX]{x1D700}_{acetonitrile}=36.6$
and
 $\unicode[STIX]{x1D700}_{water}=79.5$
.

The channel surface is uncharged (
 $\unicode[STIX]{x1D70E}=0$
), but is hydrophilic (
 $\unicode[STIX]{x1D6FE}=0.2C>0$
), resulting in the adsorption of water near the wall. Since the anions are hydrophilic, the water ‘drags’ them along, and hence the anions also effectively adsorb at the surface; see the ionic profiles in figure 2(b). Although the cations are indifferent to the local composition, they also adsorb at the surface due to electrostatics, but are distributed more broadly. The result is an electric double layer at an uncharged surface (Samin & Tsori Reference Samin and Tsori2013; Samin et al.
Reference Samin, Hod, Melamed, Gottlieb and Tsori2014), even though this layer is overall charge neutral. Figure 2 shows good agreement between the full numerical solutions and the linear theory. The numerical profiles of
 $\unicode[STIX]{x1D711}$
and
 $n^{\pm }$
decay to values that slightly differ from their bulk values due to a weak nonlinear effect induced by solvation (Samin & Tsori Reference Samin and Tsori2016). The thickness of the charged layer at the surface,
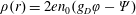 $\unicode[STIX]{x1D70C}(r)=2en_{0}(g_{D}\unicode[STIX]{x1D711}-\unicode[STIX]{x1D6F9})$
, could be estimated in experiments from the characteristic lengths
 $q_{i}^{-1}$
. These length scales are to be estimated from (3.3) given data for
 $\unicode[STIX]{x1D706}_{D}$
,
 $\unicode[STIX]{x1D709}$
and
 $g_{D}$
.
Although the fluid is overall neutral, the negative ion layer is more strongly adsorbed at the wall. When an external axial electric field is applied, this layer then serves as an effective surface charge, and the adjacent positively charged layer as an effective electric double layer. The result is an induced fluid flow, similar to EOF. The steady-state axial flow profile
 $v_{z}(r)$
is obtained by putting (3.1) and (3.2) into (2.10), and imposing the no-slip BC at
 $r=R$
and symmetry at
 $r=0$
,
 $$\begin{eqnarray}\displaystyle v_{z}(r)=\frac{\bar{\unicode[STIX]{x1D700}}E_{z}}{\unicode[STIX]{x1D6FD}e\unicode[STIX]{x1D702}}\mathop{\sum }_{i=1,2}(g_{D}b_{i}-a_{i})\frac{\text{I}_{0}(q_{i}r)-\text{I}_{0}(q_{i}R)}{(-1)^{i}(\unicode[STIX]{x1D706}_{D}q_{i})^{2}}, & & \displaystyle\end{eqnarray}$$
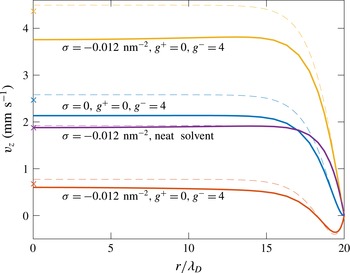
which for the parameters of figure 2 is plotted in figure 3 (labelled with
 $\unicode[STIX]{x1D70E}=0$
), showing reasonable agreement between the numerical solution and (3.6). Indeed, a flow is generated by the overall neutral but locally charged layer of fluid near the wall. Figure 3 shows that
 $v_{z}(r)$
quickly saturates to a plug-like flow of a few
 $\text{mm}~\text{s}^{-1}$
. We term this solvation-induced phenomenon solvo-osmotic flow (SOF). This flow is similar and comparable in magnitude to simple EOF, shown in figure 3 by the curve for a neat solvent with the same properties as the mixture, but near a weakly charged surface with
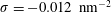 $\unicode[STIX]{x1D70E}=-0.012~\text{nm}^{-2}$
.
Solvo-osmotic flow can either combine with or work against EOF, as shown by two curves in figure 3, where we used the parameter of figure 2, but with non-zero surface charges,
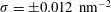 $\unicode[STIX]{x1D70E}=\pm 0.012~\text{nm}^{-2}$
. For
 $\unicode[STIX]{x1D70E}<0$
, the result is an enhanced flow with the velocity at the channel centre slightly smaller than the sum of SOF and EOF on their own. In a neat solvent, when
 $\unicode[STIX]{x1D70E}$
changes sign, one expects a simple change of sign also for the velocity. However, here SOF opposes EOF for
 $\unicode[STIX]{x1D70E}>0$
and, for the parameters of figure 3, the result is that inside most of the channel
 $v_{z}$
remains positive, but with a much smaller magnitude. There is, however, a weak backflow close to the channel wall resulting from the radial component of the body force in the fluid becoming significant in this case.
Axial velocity profiles
 $v_{z}(r)$
at an applied field of
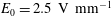 $E_{0}=2.5~\text{V}~\text{mm}^{-1}$
for several scenarios; see the labels on the figure. All other parameters are the same as in figure 2. The numerical results (solid curves) are in good agreement with the linear theory (dashed curves), where in (3.6) we used
 $E_{z}$
from the numerical calculation. Crosses were calculated using (3.8).

In the large-
 $R$
limit where
 $R\gg \unicode[STIX]{x1D706}_{D},\unicode[STIX]{x1D709}$
, the fluid moves as a plug,
 $v_{z}=\text{const}.$
, and one defines the electro-osmotic mobility of the fluid as
 $\unicode[STIX]{x1D707}_{e}=v_{z}/E_{z}$
. In this limit, we find
 $$\begin{eqnarray}\displaystyle \unicode[STIX]{x1D707}_{e}=-\frac{\bar{\unicode[STIX]{x1D700}}}{\unicode[STIX]{x1D702}}\frac{4\unicode[STIX]{x03C0}l_{B}\unicode[STIX]{x1D70E}(q_{1}^{2}+q_{2}^{2}+q_{1}q_{2}-\unicode[STIX]{x1D706}_{D}^{-2})+g_{D}\unicode[STIX]{x1D6FE}\unicode[STIX]{x1D706}_{D}^{-2}}{q_{1}q_{2}(q_{1}+q_{2})\unicode[STIX]{x1D6FD}e}. & & \displaystyle\end{eqnarray}$$
Noting that the biquadratic equation (3.3) implies
 $q_{1}q_{2}=(\unicode[STIX]{x1D709}\unicode[STIX]{x1D706}_{D})^{-1}$
and
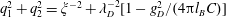 $q_{1}^{2}+q_{2}^{2}=\unicode[STIX]{x1D709}^{-2}+\unicode[STIX]{x1D706}_{D}^{-2}[1-g_{D}^{2}/(4\unicode[STIX]{x03C0}l_{B}C)]$
, and assuming that also
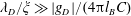 $\unicode[STIX]{x1D706}_{D}/\unicode[STIX]{x1D709}\gg |g_{D}|/(4\unicode[STIX]{x03C0}l_{B}C)$
, equation (3.7) reduces to the modified Helmholtz–Smoluchowski equation
 $$\begin{eqnarray}\displaystyle \unicode[STIX]{x1D707}_{e}=-\frac{\bar{\unicode[STIX]{x1D700}}}{\unicode[STIX]{x1D702}}\left[\left(4\unicode[STIX]{x03C0}l_{B}\unicode[STIX]{x1D706}_{D}\unicode[STIX]{x1D70E}+\frac{g_{D}\unicode[STIX]{x1D6FE}\unicode[STIX]{x1D709}}{1+\unicode[STIX]{x1D706}_{D}/\unicode[STIX]{x1D709}}\right)\frac{1}{\unicode[STIX]{x1D6FD}e}\right]. & & \displaystyle\end{eqnarray}$$
Equation (3.8) is the main result of this paper, where the term in brackets is the zeta potential
 $\unicode[STIX]{x1D701}$
, with the shear plane located at
 $r=R$
. Only the first term
 $\propto \unicode[STIX]{x1D70E}$
in (3.8) exists for a neat solvent (Keh & Tseng Reference Keh and Tseng2001). In a mixture,
 $\unicode[STIX]{x1D701}$
is determined also by the properties of the salt and surface through the second term
 $\propto g_{D}\unicode[STIX]{x1D6FE}$
in (3.8), which plays the role of an effective surface charge. The attractive feature of (3.8) is the ability to easily tune the fluid mobility through
 $g_{D}$
without making any surface modification. For example, by replacing the anion
 $\text{Cl}^{-}$
with the hydrophobic anion
 $\text{BPh}_{4}^{-}$
, we have
 $g_{D}\approx 6$
(Marcus Reference Marcus2007) (instead of
 $g_{D}=-2$
), which would then change
 $\unicode[STIX]{x1D707}_{e}$
drastically, in some cases even its sign. As we show below, the dependence of
 $\unicode[STIX]{x1D707}_{e}$
on
 $\unicode[STIX]{x1D709}$
in mixtures opens the possibility to tune the mobility also with temperature, since
 $\unicode[STIX]{x1D709}$
diverges near the mixture critical temperature. Nevertheless, we stress that
 $\unicode[STIX]{x1D709}\sim a$
for completely miscible mixtures (the
 $\unicode[STIX]{x1D712}\rightarrow 0$
limit in our theory). Hence, equation (3.8) should be applicable to mixtures in general, and, as the crosses in figure 3 confirm, it is in good agreement with the numerical results for
 $v_{z}(0)$
.
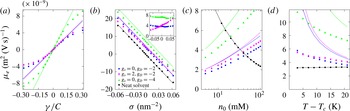
Dependence of the fluid mobility
 $\unicode[STIX]{x1D707}_{e}$
on (a) the surface–solvent coupling
 $\unicode[STIX]{x1D6FE}$
, (b) the surface charge
 $e\unicode[STIX]{x1D70E}$
, (c) the salt concentration
 $n_{0}$
and (d) the temperature
 $T$
, for three types of electrolytic mixture (see text). Symbols correspond to numerical results and lines to (3.8). In (b–d), we plot also
 $\unicode[STIX]{x1D707}_{e}$
for a neat solvent with the same physical properties as the mixtures (circles). In (a–c),
 $T=293$
K is fixed, while in (a,b,d) we keep
 $n_{0}=50$
mM fixed. For the mixture data sets in (b–d), we took
 $\unicode[STIX]{x1D6FE}=0.2C$
, and in (a,c,d), we took
 $\unicode[STIX]{x1D70E}=0$
. For the neat solvent in (c,d), we used
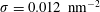 $\unicode[STIX]{x1D70E}=0.012~\text{nm}^{-2}$
. The inset of (b) shows the shift in mobility between the mixtures and the neat solvent.

In figure 4, we show the effects of varying some of the free parameters of our model on the fluid mobility, and compare numerical results (symbols) with (3.8) (lines). We compare three types of electrolytic mixture, one containing NaCl (diamonds) as before and two containing so-called antagonistic salts in which one of the ions is hydrophilic and the other hydrophobic. In order to single out the role of
 $g_{D}$
, we choose
 $g_{S}=0$
by setting
 $g^{-}=-g^{+}=2$
(squares) and 4 (triangles). Figure 4(a) shows the pure SOF mobility as a function of the surface field
 $\unicode[STIX]{x1D6FE}$
, revealing that the linear theory becomes an excellent approximation for small enough
 $|\unicode[STIX]{x1D6FE}|$
, for which
 $\unicode[STIX]{x1D711}\ll 1$
holds, and that
 $\unicode[STIX]{x1D707}_{e}$
increases with
 $\unicode[STIX]{x1D6FE}$
and
 $g_{D}$
, as expected from (3.8). It should be noticed that the sign of
 $\unicode[STIX]{x1D707}_{e}$
depends on the wetting properties of the surface. For the salts having the same solubility contrast
 $g_{D}$
but different overall solubility
 $g_{S}$
(squares and diamonds), we find significant differences in the mobility only for large enough
 $|\unicode[STIX]{x1D6FE}|$
. In this regime, the entropy decrease of the ions near the wall hinders their adsorption, leading to a reduced mobility compared with the linear theory.
In figure 4(b–d), we also contrast the electrolytic mixtures with a neat solvent (circles) having the same physical properties as the mixtures. In figure 4(b), we examine the effect of adding a fixed surface charge
 $\unicode[STIX]{x1D70E}$
. Clearly, here SOF increases the mobility compared with regular EOF in a neat solvent. The inset of figure 4(b) shows that the shift in mobility is almost constant and increases with
 $g_{D}$
for the antagonistic salts, as expected from (3.8). The mobility shift is not constant for NaCl, rather it increases with
 $\unicode[STIX]{x1D70E}$
, and the result is an asymmetry in the mobility when
 $\unicode[STIX]{x1D70E}$
changes sign, an effect that is absent in neat solvents. This effect originates from the nonlinear coupling between electrostatics and solvation (Samin & Tsori Reference Samin and Tsori2012), and is expressed by the next-order term
 $\propto g_{S}\unicode[STIX]{x1D711}\unicode[STIX]{x1D6F9}$
in the expansion of
 $\unicode[STIX]{x1D6FF}n^{\pm }$
. This contribution vanishes for our ideal antagonistic salts, but should be relevant for the majority of realistic salts. The relative shift in mobility due to SOF is significant for the weakly to moderately charged surfaces we considered here, which is a common situation for hydrophobic surfaces (Jing & Bhushan Reference Jing and Bhushan2013), or in hydrophilic surfaces near their iso-electric point. For typically highly charged hydrophilic surfaces, the contribution of SOF may be small, although this may not always be the case since both
 $g_{D}$
and
 $\unicode[STIX]{x1D6FE}$
can be much larger than the values used in this work.
In figure 4(c,d), we show the influence of the salt concentration and temperature respectively on mobility. In both panels, we compare electrolytic mixtures in an uncharged channel with a neat solvent in a channel with
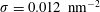 $\unicode[STIX]{x1D70E}=0.012~\text{nm}^{-2}$
. Strikingly, figure 4(c) shows that the mobility increases with
 $n_{0}$
for the mixtures, while it decreases for the neat solvent. This behaviour follows from the dependence of
 $\unicode[STIX]{x1D707}_{e}$
on
 $\unicode[STIX]{x1D706}_{D}\propto n_{0}^{-1/2}$
in (3.8). For constant
 $\unicode[STIX]{x1D70E}$
, the contribution to the surface potential from the first term in (3.8) becomes smaller when
 $\unicode[STIX]{x1D706}_{D}$
decreases, while the second term becomes larger. Figure 4(c) suggests that for
 $n_{0}\sim 100$
mM, the contribution to the mobility from solvation could be dominant when the surface is weakly charged. The SOF is enhanced by increasing the ionic strength since this leads also to increased ion adsorption at the wall, which results in a larger effective surface charge. The opposite dependence on
 $n_{0}$
between the two terms in (3.8) can therefore lead to a minimum in the mobility of mixtures at charged surfaces, which is confirmed by numerical calculations (not shown).
Figure 4(d) demonstrates that, in mixtures, temperature can be used to effectively tune the mobility, whereas its effect is much smaller in neat solvents. The reason is that the correlation length
 $\unicode[STIX]{x1D709}$
diverges as
 $T_{c}$
is approached from the one-phase region at the critical composition. In (3.8), the mobility increases with
 $\unicode[STIX]{x1D709}$
, or equivalently when the temperature is lowered towards
 $T_{c}$
, which is confirmed by the numerical results in figure 4(d). Here, the agreement with the linear theory is not as good since
 $\unicode[STIX]{x1D711}$
is no longer small when
 $T_{c}$
is approached. The numerical results show that
 $\unicode[STIX]{x1D707}_{e}$
changes by as much at 60 % in a temperature window of 20 K, whereas it essentially remains constant for the neat solvent. We did not account, however, for the change of
 $\unicode[STIX]{x1D700}$
with temperature. This effect will be at play for both mixtures and neat solvents, and should be smaller since, for example, the permittivity of water changes only by approximately 10 % in the same temperature window.
4 Conclusions
In conclusion, we have shown that the electro-osmotic mobility of mixtures contains an additional contribution due to the preferential solvation of ions. This contribution allows flow to be driven at an electrically neutral surface and the mobility to be varied significantly by changing the type of salt or the temperature. All of our results can be directly extended to the phoretic motion of particles in an applied field.
A comparison of our results with existing experiments (Valkó et al. Reference Valkó, Sirén and Riekkola1999; Grob & Steiner Reference Grob and Steiner2002) is difficult to perform at this point, because capillary electrophoresis experiments employ fused silica surfaces, which are typically highly charged and, more importantly, charge-regulating. A modification of the theory is thus required to better mimic experimental conditions. This should include the chemical dissociation equilibrium at the channel surface, and the change in the fluid viscosity with composition. For the study of transient SOF, one should also consider the asymmetry in the cation and anion diffusion constants, and their dependence on the solvent composition. Our work points towards promising possibilities for the utilization of SOF in the transport of fluids through channels and of colloids by external fields, and we hope that it will lead to systematic experiments on the electrokinetics of solvent mixtures.
Acknowledgements
We acknowledge stimulating discussions with H. B. Eral. We thank the anonymous referees for their useful comments and suggestions. R.v.R. acknowledges financial support of a Netherlands Organisation for Scientific Research (NWO) VICI grant funded by the Dutch Ministry of Education, Culture and Science (OCW). S.S. acknowledges funding from the European Union’s Horizon 2020 programme under the Marie Skłodowska-Curie grant agreement no. 656327. This work is part of the D-ITP consortium, a programme of the NWO funded by the OCW.