To save content items to your account,
please confirm that you agree to abide by our usage policies.
If this is the first time you use this feature, you will be asked to authorise Cambridge Core to connect with your account.
Find out more about saving content to .
To save content items to your Kindle, first ensure no-reply@cambridge.org
is added to your Approved Personal Document E-mail List under your Personal Document Settings
on the Manage Your Content and Devices page of your Amazon account. Then enter the ‘name’ part
of your Kindle email address below.
Find out more about saving to your Kindle.
Note you can select to save to either the @free.kindle.com or @kindle.com variations.
‘@free.kindle.com’ emails are free but can only be saved to your device when it is connected to wi-fi.
‘@kindle.com’ emails can be delivered even when you are not connected to wi-fi, but note that service fees apply.
A gas flow in a two-dimensional square cavity driven by a lid moving parallel to itself in the cavity plane is considered. The Stokes equations under the no-slip condition predict an infinite series of corner vortices near the lower corners, flow velocity with direction-dependent limiting values at the upper corners, and stress and pressure that diverge inversely with the distance from the upper corners. We examine how these properties are modified or retained within the kinetic theory of gases. The steady behaviour of the gas is numerically investigated based on the linearised Bhatnagar–Gross–Krook kinetic equation and the diffuse reflection boundary condition. No corner vortices are observed for $k \ge 0.03$, where $k = (\sqrt {\pi }/2)\textit{Kn}$ with $\textit{Kn}$ being the Knudsen number. A first corner vortex is observed at $k = 0.02$ and for smaller $k$ examined. A second one is observed at $k = 0.001$ and for smaller $k$ examined. Numerical results imply that their number is finite for each Knudsen number. Macroscopic quantities, including the flow velocity, exhibit direction-dependent limiting values also within kinetic theory. For moderate to large Knudsen numbers, such direction dependence manifests itself even at the lower corners. The stress and pressure vary inversely with the distance from the upper corners up to a few tens of mean free paths. Their increase is suppressed at positions closer to the corners and they remain bounded. Consequently, the total forces acting on the lid and side cavity walls are also finite, and they exhibit an unconventional scaling $\textit{Kn} \ln \textit{Kn}$ for small $\textit{Kn}$.
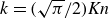
We present a new kinetic model and its lattice Boltzmann realisation for the simulation of compressible, non-ideal fluid flows. The method employs first-neighbour lattices and introduces a consistent set of correction terms constructed via quasi-equilibrium attractors, ensuring positive-definite and Galilean-invariant Navier–Stokes dissipation rates. This construction circumvents the need for extended stencils or ad hoc regularisation, while maintaining numerical stability and thermodynamic consistency across a broad range of flow regimes. The resulting model accurately reproduces both the Euler and Navier–Stokes hydrodynamic limits. As a stringent validation, we demonstrate, for the first time within a lattice Boltzmann framework, quantitatively accurate simulations of shock–drop interactions at Mach numbers up to 1.47. The proposed approach thus extends the applicability of lattice Boltzmann methods to high-speed, non-ideal compressible flows with a minimal kinetic stencil.
Droplet ripening is conventionally modelled by macroscopic interfacial theories, which inherently neglect the intrinsic microscopic structures of the liquid–vapour system, specifically the finite diffuse interface and the adjacent Knudsen layer. Resolving these structures is critical, as they govern mass transfer rates in regimes where interfacial transport kinetics dominate over bulk diffusion, particularly in micro- and nanodroplet interactions. To elucidate the underlying fundamental physical mechanisms, we investigate droplet ripening in a single-component liquid–vapour system using the Enskog–Vlasov kinetic model. This approach naturally captures the full spatial structure of both the diffuse interface and the Knudsen layer. We first focus on two-droplet interactions, deriving a theoretical growth rate based on the intrinsic imbalance of molecular fluxes at the interface. Numerical simulations quantitatively validate this prediction, confirming that the droplet-radius change rate scales linearly with the thermodynamic driving force. However, a distinct deviation from this linear behaviour emerges in the late stage of ripening. We attribute this breakdown to the overlap of diffuse interfaces, which reduces the liquid density and induces thermodynamic destabilisation. Finally, our simulations of three-droplet systems confirm the universality of the proposed theoretical model. These findings provide a microscopic understanding of ripening and help bridge nanoscale dynamics with macroscopic coarsening theories.
This study investigates the non-equilibrium transport characteristics of pressure-driven microscale and rarefied Poiseuille flows based on analytical modelling and the direct simulation Monte Carlo (DSMC) method. A closed-form analytical solution for the cross-stream pressure and viscous stress distributions is derived from the second-order nonlinear coupled constitutive relation model. This solution reveals a compact tangent-function form, demonstrating that the convex pressure profiles observed in rarefied flows – unaccounted for by classical Navier–Stokes–Fourier theory – are driven by the coupling of kinematic stress constraints and geometric confinement. Complementing this theory, an extensive set of DSMC simulations is performed across a wide range of Knudsen numbers (up to 13.23), Mach numbers (up to 0.868) and channel aspect ratios. These simulations analyse the effects of rarefaction and pressure ratio, with a special emphasis on entropy and entropy generation. A kinetic decomposition of the heat flux and an energy-budget analysis demonstrate a pronounced non-Fourier regime in highly rarefied channels, in which the streamwise heat flux reverses sign and transports energy upstream and from colder core regions towards the hotter inlet, extending linear Onsager-based predictions to nonlinear, pressure-driven microchannel flows at moderate Mach number. This anomaly arises not from anomalous diffusion but from stress–work coupling between anisotropic normal stresses and velocity gradients that force the heat-flux divergence to balance expansion work when convective cooling collapses. Our findings highlight the importance of non-equilibrium constitutive models to accurately predict irreversible processes in microscale and rarefied gas flows inside channels, establishing the analytical and computational results as reference data.
Non-equilibrium evaporative flows play a central role in many nanoporous membrane technologies, where transport of fluids is confined by solid surfaces at the nanoscale. In this work, we propose a molecular kinetic model that consistently resolves the coupled interactions among vapour, liquid and solid surfaces in such flows. As a direct consequence of this bottom-up approach, the liquid–vapour, liquid–solid and vapour–solid interfaces form autonomously, and the effects of non-equilibrium and real fluids can be captured simultaneously, which does not need empirical models depending on ad hoc parameters such as the evaporation/condensation coefficients and contact angle. Accuracy of the model improves further by including the soft-collision effect in the pair correlation function and applying a temperature-dependent correction to the mean-field Vlasov term, as validated against the experimental data and the molecular dynamics simulations. Furthermore, when applied to unsteady, evaporation-driven liquid–vapour flows, the model reveals distinct dynamics due to surface wettability: the hydrophilic surfaces exhibit phenomena such as liquid meniscus breakage and enhanced evaporation flux, whereas the hydrophobic surfaces lead to disappearance of liquid droplets. These findings highlight the potential of the proposed molecular kinetic model as a powerful design tool for next-generation nano-technologies that leverage nano-confined phase change.
Kinetic theory offers a promising alternative to conventional turbulence modelling by providing a mesoscopic perspective that naturally captures non-equilibrium physics such as non-Newtonian effects. In this work, we present an extension and theoretical analysis of the kinetic model for incompressible turbulent flows developed by Chen et al. (Atmosphere, 2023, vol. 14(7), p. 1109), constructed for unbounded flows. The first extension is to reselect a relaxation time such that the turbulent transport coefficients are obtained consistently and better align with well-established turbulence theory. The Chapman–Enskog (CE) analysis of the kinetic model reproduces the linear eddy-viscosity and gradient diffusion models for Reynolds stress and turbulent kinetic energy flux at the first order, and yields nonlinear eddy-viscosity and closure models at the second order. In particular, a previously unreported CE solution for turbulent kinetic energy flux is obtained. The second extension is to enable the model for wall-bounded turbulent flows with preserved near-wall asymptotic behaviours. This involves developing a low-Reynolds-number model incorporating wall damping effects and viscous diffusion, with boundary conditions enabling both viscous sublayer resolution and wall function application. Comprehensive validation against experimental and direct numerical simulation data for turbulent Couette flow demonstrates excellent agreement in predicting mean velocity profiles, skin friction coefficients and Reynolds shear-stress distributions, although the near-wall-normal stress anisotropy is underestimated. The results show that averaged turbulent flow behaves similarly to rarefied-gas flow at finite Knudsen number, capturing non-Newtonian effects beyond linear eddy-viscosity models. This kinetic model provides a physics-based foundation for turbulence modelling with reduced empirical dependence.
We derive a self-consistent hydrodynamic theory of coupled binary fluid–surfactant systems from the underlying microscopic physics using Rayleigh’s variational principle. At the microscopic level, surfactant molecules are modelled as dumbbells that exert forces and torques on the fluid and interface while undergoing Brownian motion. We obtain the overdamped stochastic dynamics of these particles from a Rayleighian dissipation functional, which we then coarse-grain to derive a set of continuum equations governing the surfactant concentration, orientation, fluid density and velocity. This approach introduces a polarisation field $\boldsymbol{p}(\boldsymbol{r},t)$, representing the average orientation of surfactants, which plays a central role in suppressing droplet coalescence. The remaining hydrodynamic equations are consistently obtained from a mesoscopic free energy functional. The resulting model accurately captures key surfactant phenomena, including surface tension reduction and droplet stabilisation, as confirmed by both perturbation theory and numerical simulations, and is thermodynamically consistent with both the Gibbs adsorption isotherm and Henry’s law for adsorbed surfactant concentration.
The classical problem of steady rarefied gas flow past an infinitely thin circular disk is revisited, with particular emphasis on the gas behaviour near the disk edge. The uniform flow is assumed to be perpendicular to the disk surface. An integral equation for the velocity distribution function, derived from the linearised Bhatnagar–Gross–Krook model of the Boltzmann equation and subject to diffuse reflection boundary conditions, is solved numerically. The numerical method fully accounts for the discontinuity in the velocity distribution function that arises due to the presence of the edge. It is found that a kinetic boundary layer forms near the disk edge, extending over several mean free paths, and that its magnitude scales as $\textit{Kn}^{1/2}$ as the Knudsen number $\textit{Kn}$ (defined with respect to the disk radius) tends to zero. A thermal polarisation effect, previously studied for spherical geometries, is also observed in the disk case, with a more pronounced manifestation near the edge that exhibits the same $\textit{Kn}^{1/2}$ scaling. The drag force acting on the disk is computed over a wide range of Knudsen numbers and shows good agreement with existing results for a hard-sphere gas and in the near-free-molecular regime.
Near-space hypersonic vehicles encounter significant rarefaction effects during the flight through the atmosphere, causing the classical Navier–Stokes–Fourier (NSF) equations to break down and posing challenges for the evaluation of surface drag and heat flux. In this paper, the nonlinear momentum and heat transfer in a hypersonic transitional boundary layer are analysed based on the generalized hydrodynamic equations (GHE), and the generality of the derived formulae is also discussed. The leading transport relations are obtained by estimating the relative orders of the various terms in GHE according to the hypersonic flow and boundary-layer requirements. Local non-equilibrium parameters characterising the shear non-equilibrium effect ($K_\sigma$) and thermal-gradient non-equilibrium effect ($K_q$) are introduced, and a set of correlation formulae for local surface pressure, shear stress and heat flux are proposed as corrections to continuum-based solutions. The correction function depends only on the non-equilibrium parameters $K_\sigma$ and $K_q$, and the continuous solutions can be either analytical formulae or NSF simulation results. This enables us to predict the surface aerothermodynamics with enhanced accuracy while still using the solutions of the NSF equations. The proposed formulae are carefully verified by comparing with direct simulation Monte Carlo (DSMC) results of different hypersonic rarefied flows, including flat-plate, sharp-wedge, cylinder and blunt-cone flows, and partial experimental data are also given. The results demonstrate that the proposed formulae can significantly enhance the accuracy of the continuum-based solutions, and show good agreement with DSMC simulations and experimental measurements in the near-continuum regime.
New experimental results on gas flow through a long tube in the viscous, slip and transitional regimes are presented, obtained using an improved constant-volume measurement technique. This method is based on measuring the pressure variation in the inlet tank while the outlet tank is evacuated to a low pressure. Experimental pressure data for helium, neon, argon, nitrogen, krypton and xenon are used to extract the Poiseuille coefficient through a newly developed methodology. The obtained values show good agreement with theoretical predictions. Additionally, the velocity slip coefficient is also extracted from the same pressure data for all tested gases.
Kinetic theory is summarized as a mechanistic approach to thermodynamics, including the equilibrium state equation of an ideal gas and a phenomenological approach to its transport properties. The Boltzmann model of the ideal gas is described by the evolution equation of its distribution function in molecular space. The H-theorem is proved for both the uniform and nonuniform cases. The theorem of additive invariants allows to approach a fundamental formulation of hydrodynamic equations for both the ideal situation of an inviscid flow and for the more interesting case of a viscous flow.
Take anything in the universe, put it in a box, and heat it up. Regardless of what you start with, the motion of the substance will be described by the equations of fluid mechanics. This remarkable universality is the reason why fluid mechanics is important. The key equation of fluid mechanics is the Navier-Stokes equation. This textbook starts with the basics of fluid flows, building to the Navier-Stokes equation while explaining the physics behind the various terms and exploring the astonishingly rich landscape of solutions. The book then progresses to more advanced topics, including waves, fluid instabilities, and turbulence, before concluding by turning inwards and describing the atomic constituents of fluids. It introduces ideas of kinetic theory, including the Boltzmann equation, to explain why the collective motion of 1023 atoms is, under the right circumstances, always governed by the laws of fluid mechanics.
Statistical mechanics is hugely successful when applied to physical systems at thermodynamic equilibrium; however, most natural phenomena occur in nonequilibrium conditions and more sophisticated techniques are required to address this increased complexity. This second edition presents a comprehensive overview of nonequilibrium statistical physics, covering essential topics such as Langevin equations, Lévy processes, fluctuation relations, transport theory, directed percolation, kinetic roughening, and pattern formation. The first part of the book introduces the underlying theory of nonequilibrium physics, the second part develops key aspects of nonequilibrium phase transitions, and the final part covers modern applications. A pedagogical approach has been adopted for the benefit of graduate students and instructors, with clear language and detailed figures used to explain the relevant models and experimental results. With the inclusion of original material and organizational changes throughout the book, this updated edition will be an essential guide for graduate students and researchers in nonequilibrium thermodynamics.
This chapter begins with a formal definition of a fluid (what it means to be a continuum rather than an ensemble of particles) followed by a review of kinetic theory of gases where the connections between pressure and particle momentum and between specific energy (temperature) and average particle kinetic energy are made. A distinction is made between extensive and intensive variables, from which the Theorem of Hydrodynamics is postulated and proven. From this theorem, the basic equations of ideal hydrodynamics (zero-field limit of MHD) are derived including continuity, total energy equation, and the momentum equation. Alternate equations of HD such as the internal energy, pressure, and Euler’s equations are also introduced. The equations of HD are then assembled into two sets–conservative and primitive–with the distinction between the two explained.
We have established a novel molecular kinetic model that addresses fundamental challenges in the non-equilibrium transport of nanoscale confined fluids, such as rarefaction and fluid inhomogeneities, which are crucial to a range of scientific and engineering fields. The proposed model explicitly considers fluid–solid molecular interactions in the transport equations, eliminating the reliance on predefined boundary conditions. By consistently accounting for molecular interactions between fluids and solids, the unified model captures both intrinsic and apparent non-hydrodynamic effects, as well as real fluid behaviours. Rigorous comparisons with molecular dynamics simulations demonstrate that the present model accurately predicts unique features of strongly inhomogeneous fluid flows, including fluid adsorption, solvation force, velocity slip and temperature jump. Therefore, this mesoscopic model bridges the gap between molecular-scale dynamics and macroscopic hydrodynamics, enabling a practical simulation tool for nanoscale surface-confined flows. Moreover, it offers valuable insights into the molecular mechanisms underlying anomalous transport phenomena observed in confined flows, such as the disappearance and re-emergence of the Knudsen minimum.
This study presents a novel approach for constructing turbulence models using the kinetic Fokker–Planck equation. By leveraging the inherent similarities between Brownian motion and turbulent dynamics, we formulate a Fokker–Planck equation tailored for turbulence at the hydrodynamic level. In this model, turbulent energy plays a role analogous to temperature in molecular thermodynamics, and the large-scale structures are characterised by a turbulent relaxation time. This model aligns with the framework of Pope’s generalised Langevin model, with the first moment recovering the Reynolds-averaged Navier–Stokes (RANS) equations, and the second moment yielding a partially modelled Reynolds stress transport equation. Utilising the Chapman–Enskog expansion, we derive asymptotic solutions for this turbulent Fokker–Planck equation. With an appropriate choice of relaxation time, we obtain a linear eddy viscosity model at first order, and a quadratic Reynolds stress constitutive relationship at second order. Comparative analysis of the coefficients of the quadratic expression with typical nonlinear viscosity models reveals qualitative consistency. To further validate this kinetic-based nonlinear viscosity model, we integrate it as a RANS model within computational fluid dynamics codes, and calculate three typical cases. The results demonstrate that this quadratic eddy viscosity model outperforms the linear model and shows comparability to a cubic model for two-dimensional flows, without the introduction of ad hoc parameters in the Reynolds stress constitutive relationship.
A new lattice Boltzmann model (LBM) is presented to describe chemically reacting multicomponent fluid flow in homogenised porous media. In this work, towards further generalising the multicomponent reactive lattice Boltzmann model, we propose a formulation which is capable of performing reactive multicomponent flow computation in porous media at the representative elementary volume (REV) scale. To that end, the submodel responsible for interspecies diffusion has been upgraded to include Knudsen diffusion, whereas the kinetic equations for the species, the momentum and the energy have been rewritten to accommodate the effects of volume fraction of porous media through careful choice of the equilibrium distribution functions. Verification of the mesoscale kinetic system of equations by a Chapman–Enskog analysis reveals that at the macroscopic scale, the homogenised Navier–Stokes equations for compressible multicomponent reactive flows are recovered. The dusty gas model (DGM) capability hence formulated is validated over a wide pressure range by comparison of experimental flow rates of component species counter diffusing through capillary tubes. Next, for developing a capability to compute heterogeneous reactions, source terms for maintaining energy and mass balance across the fluid phase species and the surface adsorbed phase species are proposed. The complete model is then used to perform detailed chemistry simulations in porous electrodes of a solid oxide fuel cell (SOFC), thereby predicting polarisation curves which are of practical interest.
We develop the time-dependent regularised 13-moment equations for general elastic collision models under the linear regime. Detailed derivation shows the proposed equations have super-Burnett order for small Knudsen numbers, and the moment equations enjoy a symmetric structure. A new modification of Onsager boundary conditions is proposed to ensure stability as well as the removal of undesired boundary layers. Numerical examples of one-dimensional channel flows is conducted to verified our model.
Starting from the coupled Boltzmann–Enskog (BE) kinetic equations for a two-particle system consisting of hard spheres, a hyperbolic two-fluid model for binary, hard-sphere mixtures was derived in Fox (2019, J. Fluid Mech.877, 282). In addition to spatial transport, the BE kinetic equations account for particle–particle collisions, using an elastic hard-sphere collision model, and the Archimedes (buoyancy) force due to spatial gradients of the pressure in each phase, as well as other forces involving spatial gradients. The ideal-fluid–particle limit of this model is found by letting one of the particle diameters go to zero while the other remains finite. The resulting two-fluid model has closed terms for the spatial fluxes and momentum exchange due to the excluded volume occupied by the particles, e.g. a momentum-exchange term $\boldsymbol {F}_{\!\!fp}$ that depends on gradients of the fluid density $\rho _f$, fluid velocity $\boldsymbol{u}_{f}$ and fluid pressure $p_f$. In Zhang et al. (2006, Phy. Rev. Lett.97, 048301), the corresponding unclosed momentum-exchange term depends on the divergence of an unknown particle–fluid–particle (pfp) stress (or pressure) tensor. Here, it is shown that the pfp-pressure tensor ${\unicode{x1D64B}}_{\!pfp}$ can be found in closed form from the expression for $\boldsymbol {F}_{\!\!fp}$ derived in Fox (2019, J. Fluid Mech.877, 282). Remarkably, using this expression for ${\unicode{x1D64B}}_{\!pfp}$ ensures that the two-fluid model for ideal-fluid–particle flow is well posed for all fluid-to-particle material-density ratios $Z = \rho _f / \rho _p$.
Recent years have seen the emergence of new technologies that exploit nanoscale evaporation, ranging from nanoporous membranes for distillation to evaporative cooling in electronics. Despite the increasing depth of fundamental knowledge, there is still a lack of simulation tools capable of capturing the underlying non-equilibrium liquid–vapour phase changes that are critical to these and other such technologies. This work presents a molecular kinetic theory model capable of describing the entire flow field, i.e. the liquid and vapour phases and their interface, while striking a balance between accuracy and computational efficiency. In particular, unlike previous kinetic models based on the isothermal assumption, the proposed model can capture the temperature variations that occur during the evaporation process, yet does not require the computational resources of more complicated mean-field kinetic approaches. We assess the present kinetic model in three test cases: liquid–vapour equilibrium, evaporation into near-vacuum condition, and evaporation into vapour. The results agree well with benchmark solutions, while reducing the simulation time by almost two orders of magnitude on average in the cases studied. The results therefore suggest that this work is a stepping stone towards the development of an accurate and efficient computational approach to optimising the next generation of nanotechnologies based on nanoscale evaporation.